
2026
David Mendive-Tapia, Christoph Schran, Banshi Das, Fabien Gatti, Markus Schröder, Dominik Marx, Oriol Vendrell
Deciphering the infrared spectrum of the hydrated proton using full-dimensional quantum dynamics Journal Article
In: Nat. Chem., 2026.
Abstract | Links | BibTeX | Tags: Computational Chemistry, Quantum Dynamics, Water
@article{MendiveTapia2026/10.1038/s41557-026-02209-3,

title = {Deciphering the infrared spectrum of the hydrated proton using full-dimensional quantum dynamics},

author = {David Mendive-Tapia and Christoph Schran and Banshi Das and Fabien Gatti and Markus Schröder and Dominik Marx and Oriol Vendrell},

url = {https://www.nature.com/articles/s41557-026-02209-3},

doi = {10.1038/s41557-026-02209-3},

year  = {2026},

date = {2026-07-27},

journal = {Nat. Chem.},

abstract = {The nature of the aqueous proton has been traditionally interpreted through two limiting structural motifs: the Zundel and Eigen cations. However, experimental infrared (IR) spectra of the solvated proton reveal a far more dynamic character, as evidenced by distinct intensity modulations within the characteristic continuum absorption band. In fact, recent ultrafast two-dimensional IR spectroscopy suggests that solvation-induced structural distortions around H2O⋯H+⋯OH2 motifs critically shape the IR response. Here we investigate the role of such asymmetry through full-dimensional quantum dynamics simulations of the extended Zundel complex H⁺(H₂O)₆, which structurally encompasses both Zundel and Eigen motifs. Systematic removal of one water molecule from the second solvation shell gradually introduces deviations from the perfectly symmetric Zundel-like complex towards Eigen-like spectral features. These results provide a direct map between the asymmetric solvation environment and the structural response of the first and second solvation shells of the aqueous proton, offering a structural and dynamical basis for understanding how this asymmetry governs proton mobility in aqueous environments.},

keywords = {Computational Chemistry, Quantum Dynamics, Water},

pubstate = {published},

tppubtype = {article}

}
Marcella Iannuzzi, Jan Wilhelm, Frederick Stein, Augustin Bussy, Hossam Elgabarty, Dorothea Golze, Anna-Sophia Hehn, Maximilian Graml, Stepan Marek, Beliz Sertcan Gökmen, Christoph Schran, Harald Forbert, Rustam Z. Khaliullin, Anton Kozhevnikov, Mathieu Taillefumier, Rocco Meli, Vladimir V. Rybkin, Martin Brehm, Robert Schade, Ole Schütt, Johann V. Pototschnig, Hossein Mirhosseini, Andreas Knüpfer, Dominik Marx, Matthias Krack, Jürg Hutter, Thomas D. Kühne
The CP2K program package made simple Journal Article
In: J. Phys. Chem. B, vol. 130, iss. 4, pp. 1237–1310, 2026.
Abstract | Links | BibTeX | Tags: Ab-Initio Molecular Dynamics (AIMD), Density Functional Theory (DFT), Quantum Dynamics
@article{Iannuzzi2026/10.1021/acs.jpcb.5c05851,

title = {The CP2K program package made simple},

author = {Marcella Iannuzzi and Jan Wilhelm and Frederick Stein and Augustin Bussy and Hossam Elgabarty and Dorothea Golze and Anna-Sophia Hehn and Maximilian Graml and Stepan Marek and Beliz Sertcan Gökmen and Christoph Schran and Harald Forbert and Rustam Z. Khaliullin and Anton Kozhevnikov and Mathieu Taillefumier and Rocco Meli and Vladimir V. Rybkin and Martin Brehm and Robert Schade and Ole Schütt and Johann V. Pototschnig and Hossein Mirhosseini and Andreas Knüpfer and Dominik Marx and Matthias Krack and Jürg Hutter and Thomas D. Kühne},

url = {https://pubs.acs.org/doi/full/10.1021/acs.jpcb.5c05851},

doi = {10.1021/acs.jpcb.5c05851},

year  = {2026},

date = {2026-01-15},

urldate = {2026-01-15},

journal = {J. Phys. Chem. B},

volume = {130},

issue = {4},

pages = {1237–1310},

abstract = {CP2K is a versatile open-source software package for simulations across a wide range of atomistic systems, from isolated molecules in the gas phase to low-dimensional functional materials and interfaces, as well as highly symmetric crystalline solids, disordered amorphous glasses, and weakly interacting soft-matter systems in the liquid state and in solution. This review highlights CP2K's capabilities for computing both static and dynamical properties using quantum-mechanical and classical simulation methods. In contrast to the accompanying theory and code paper [J. Chem. Phys. 152, 194103 (2020)], the focus here is on the practical usage and applications of CP2K, with underlying theoretical concepts introduced only as needed.},

keywords = {Ab-Initio Molecular Dynamics (AIMD), Density Functional Theory (DFT), Quantum Dynamics},

pubstate = {published},

tppubtype = {article}

}
2024
Richard Beckmann, Christoph Schran, Fabien Brieuc, Dominik Marx
Theoretical infrared spectroscopy of protonated methane isotopologues Journal Article
In: Phys. Chem. Chem. Phys., vol. 26, iss. 35, pp. 22846-22852, 2024.
Abstract | Links | BibTeX | Tags: Coupled Cluster (CC), Quantum Dynamics, Spectra
@article{Beckmann2024/10.1039/D4CP02295E,

title = {Theoretical infrared spectroscopy of protonated methane isotopologues},

author = {Richard Beckmann and Christoph Schran and Fabien Brieuc and Dominik Marx},

url = {http://dx.doi.org/10.1039/D4CP02295E},

doi = {10.1039/D4CP02295E},

year  = {2024},

date = {2024-08-13},

urldate = {2024-08-13},

journal = {Phys. Chem. Chem. Phys.},

volume = {26},

issue = {35},

pages = {22846-22852},

publisher = {The Royal Society of Chemistry},

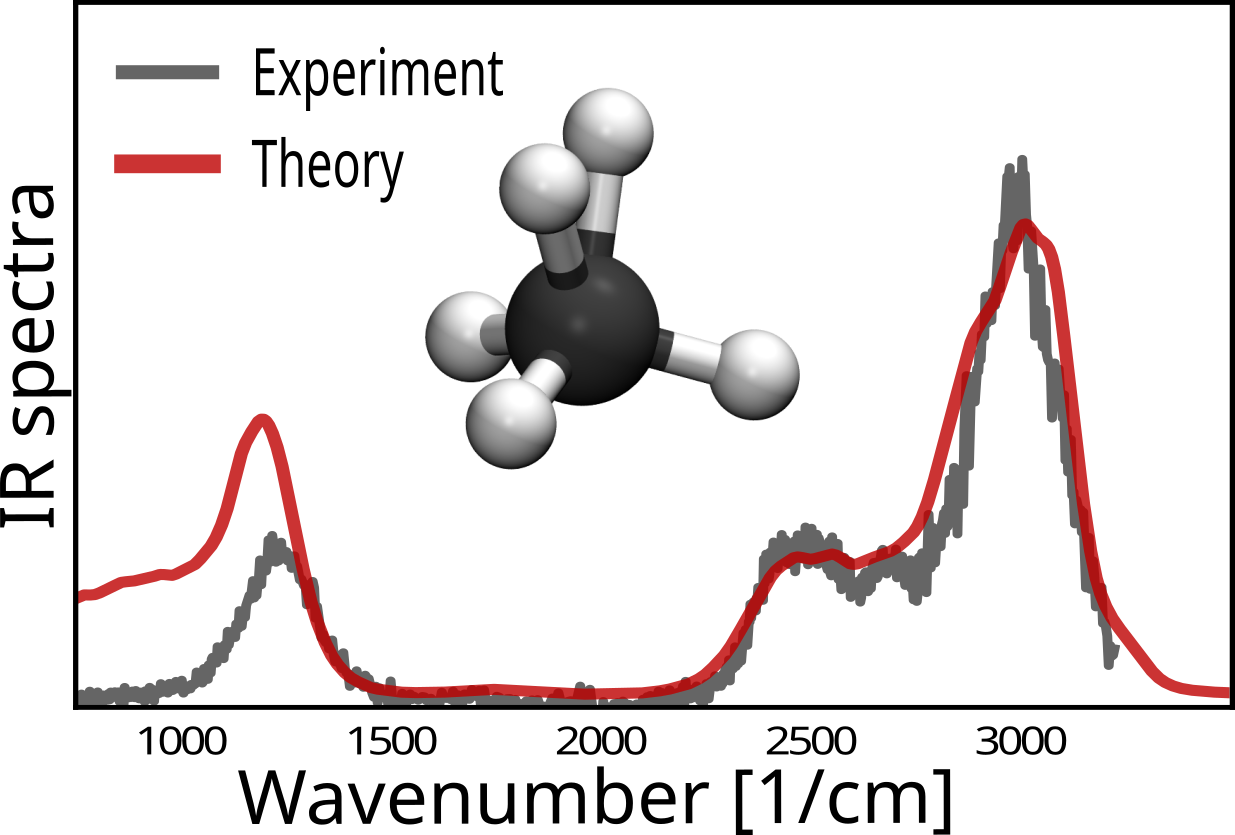
abstract = {The vibrational spectroscopy of protonated methane and its mixed hydrogen/deuterium isotopologues remains a challenge to both experimental and computational spectroscopy due to the iconic floppiness of CH5+. Here, we compute the finite-temperature broadband infrared spectra of CH5+ and all its isotopologues, i.e. CHnD5−n+ up to CD5+, from path integral molecular dynamics in conjunction with interactions and dipoles computed consistently at CCSD(T) coupled cluster accuracy. The potential energy and dipole moment surfaces have been accurately represented in full dimensionality in terms of high-dimensional neural networks. The resulting computational efficiency allows us to establish CCSD(T) accuracy at the level of converged path integral simulations. For all six isotopologues, the computed broadband spectra compare very favorably to the available experimental broadband spectra obtained from laser induced reactions action vibrational spectroscopy. The current approach is found to consistently and significantly improve on previous calculations of these broadband vibrational spectra and defines the new cutting-edge for what has been dubbed the “enfant terrible” of molecular spectroscopy in view of its pronounced large-amplitude motion that involves all intramolecular degrees of freedom.},

keywords = {Coupled Cluster (CC), Quantum Dynamics, Spectra},

pubstate = {published},

tppubtype = {article}

}
2023
Irén Simkó, Christoph Schran, Fabien Brieuc, Csaba Fábri, Oskar Asvany, Stephan Schlemmer, Dominik Marx, Attila G. Császár
Quantum nuclear delocalization and its rovibrational fingerprints Journal Article
In: Angewandte Chemie International Edition, vol. 62, no. 41, pp. e202306744, 2023.
Abstract | Links | BibTeX | Tags: Nuclear Quantum Effects (NQEs), Path Integral Molecular Dynamics (PIMD), Quantum Dynamics
@article{Simko/10.1002/anie.202306744,

title = {Quantum nuclear delocalization and its rovibrational fingerprints},

author = {Irén Simkó and Christoph Schran and Fabien Brieuc and Csaba Fábri and Oskar Asvany and Stephan Schlemmer and Dominik Marx and Attila G. Császár},

url = {https://onlinelibrary.wiley.com/doi/abs/10.1002/anie.202306744},

doi = {10.1002/anie.202306744},

year  = {2023},

date = {2023-01-01},

urldate = {2023-01-01},

journal = {Angewandte Chemie International Edition},

volume = {62},

number = {41},

pages = {e202306744},

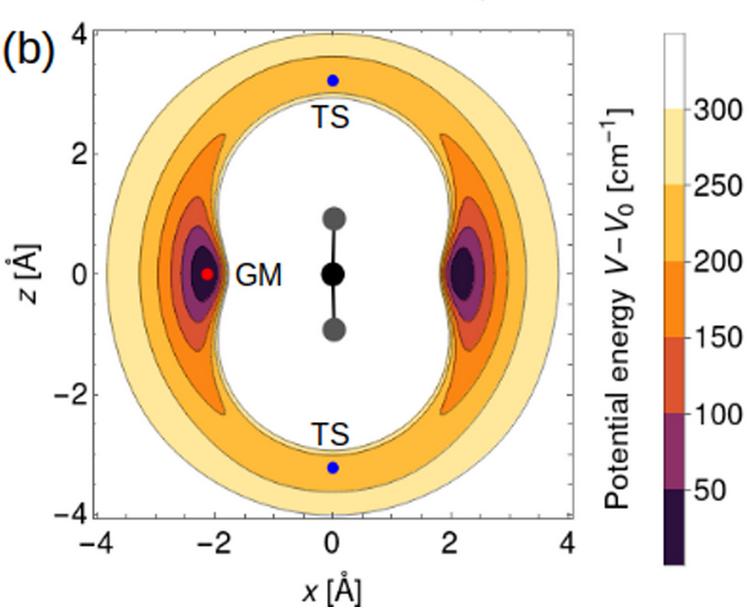
abstract = {Abstract Quantum mechanics dictates that nuclei must undergo some delocalization. In this work, emergence of quantum nuclear delocalization and its rovibrational fingerprints are discussed for the case of the van der Waals complex . The equilibrium structure of is planar and T-shaped, one He atom solvating the quasi-linear He−H+−He core. The dynamical structure of , in all of its bound states, is fundamentally different. As revealed by spatial distribution functions and nuclear densities, during the vibrations of the molecule the solvating He is not restricted to be in the plane defined by the instantaneously bent chomophore, but freely orbits the central proton, forming a three-dimensional torus around the chromophore. This quantum delocalization is observed for all vibrational states, the type of vibrational excitation being reflected in the topology of the nodal surfaces in the nuclear densities, showing, for example, that intramolecular bending involves excitation along the circumference of the torus.},

keywords = {Nuclear Quantum Effects (NQEs), Path Integral Molecular Dynamics (PIMD), Quantum Dynamics},

pubstate = {published},

tppubtype = {article}

}
2022
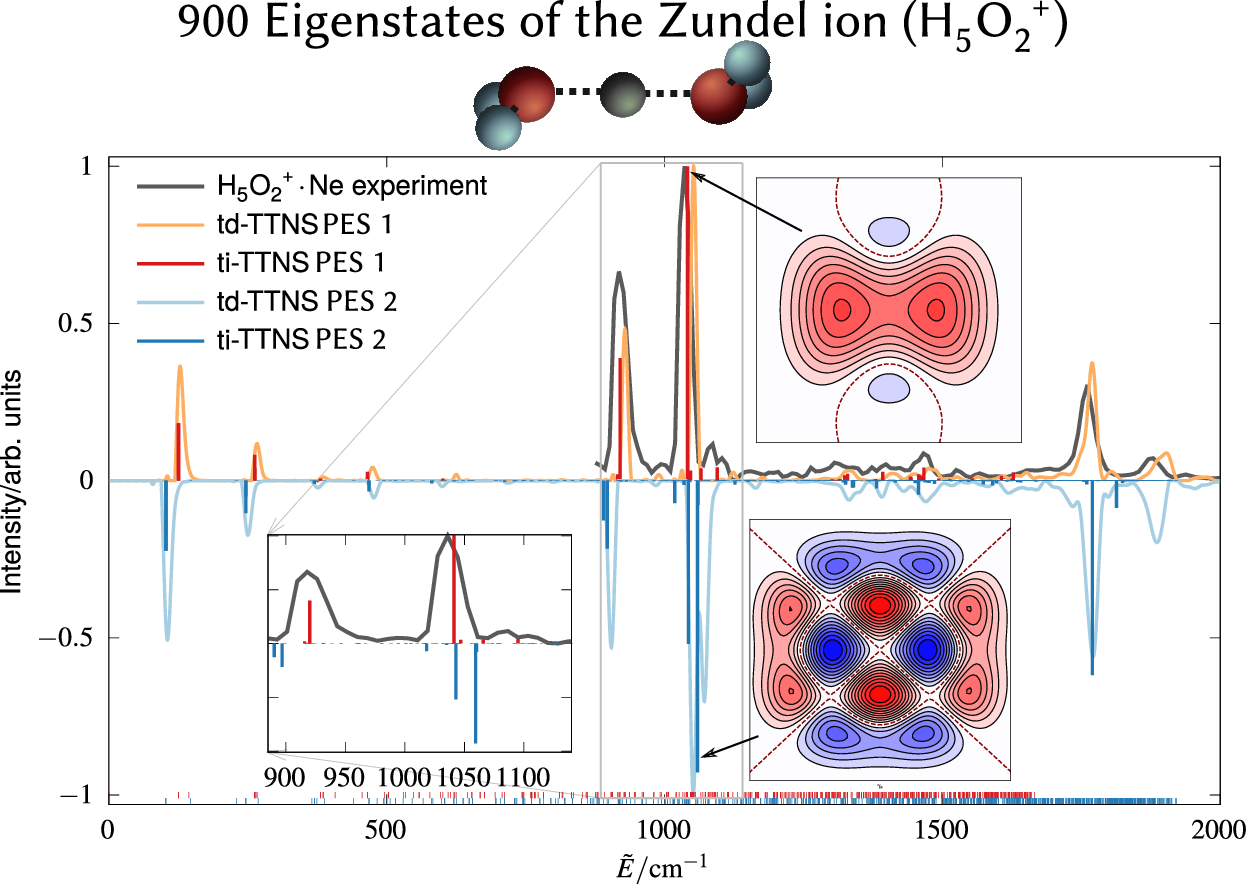
Henrik R Larsson, Markus Schröder, Richard Beckmann, Fabien Brieuc, Christoph Schran, Dominik Marx, Oriol Vendrell
State-resolved infrared spectrum of the protonated water dimer: Revisiting the characteristic proton transfer doublet peak Journal Article
In: Chem. Sci., 2022, ISSN: 2041-6539.
Abstract | Links | BibTeX | Tags: Quantum Dynamics, Spectra, Water
@article{Larsson2022/10.1039/D2SC03189B,

title = {State-resolved infrared spectrum of the protonated water dimer: Revisiting the characteristic proton transfer doublet peak},

author = {Henrik R Larsson and Markus Schröder and Richard Beckmann and Fabien Brieuc and Christoph Schran and Dominik Marx and Oriol Vendrell},

doi = {10.1039/D2SC03189B},

issn = {2041-6539},

year  = {2022},

date = {2022-08-01},

urldate = {2022-08-01},

journal = {Chem. Sci.},

publisher = {The Royal Society of Chemistry},

abstract = {The infrared (IR) spectra of protonated water clusters encode precise information on the dynamics and structure of the hydrated proton. However, the strong anharmonic coupling and quantum effects of these elusive species remain puzzling up to the present day. Here, we report unequivocal evidence that the interplay between the proton transfer and the water wagging motions in the protonated water dimer (Zundel ion) giving rise to the characteristic doublet peak is both more complex and more sensitive to subtle energetic changes than previously thought. In particular, hitherto overlooked low-intensity satellite peaks in the experimental spectrum are now unveiled and mechanistically assigned. Our findings rely on the comparison of IR spectra obtained using two highly accurate potential energy surfaces in conjunction with highly accurate state-resolved quantum simulations. We demonstrate that these high-accuracy simulations are important for providing definite assignments of the complex IR signals of fluxional molecules.},

keywords = {Quantum Dynamics, Spectra, Water},

pubstate = {published},

tppubtype = {article}

}