
2026

Darren Wayne Lim, Xavier R. Advincula, William C. Witt, Angelos Michaelides, Fabian L. Thiemann, Christoph Schran
Strain-dependent wetting of graphene Journal Article
In: Chem. Sci., 2026.
Abstract | Links | BibTeX | Tags: Machine Learning Potentials, Two-dimensional Materials, Water, Water at Interfaces
@article{Lim2026/10.1039/d6sc05377g,

title = {Strain-dependent wetting of graphene},

author = {Darren Wayne Lim and Xavier R. Advincula and William C. Witt and Angelos Michaelides and Fabian L. Thiemann and Christoph Schran},

url = {https://doi.org/10.1039/d6sc05377g},

doi = {10.1039/d6sc05377g},

year  = {2026},

date = {2026-10-20},

urldate = {2026-10-20},

journal = {Chem. Sci.},

abstract = {Understanding how water wets graphene is critical for predicting and controlling its behaviour in nanofluidic, sensing, and energy applications. A key measure of wetting is the contact angle made by a liquid droplet against the surface, yet experimental measurements for graphene span a wide range, with no consensus for free-standing graphene. Here, we use a machine learning potential with ab initio accuracy to provide an atomistic first-principles prediction for this unsolved problem, finding a weakly hydrophilic contact angle of 72.1 ± 1.5°. More importantly, we unveil that graphene's wetting properties are highly sensitive to mechanical strain: tensile strain makes graphene significantly less hydrophilic, while compressive strain induces coherent ripples around the droplet, resulting in pronounced anisotropic wetting and contact angle hysteresis. We show that there is a strong coupling between the three-phase contact line and the intrinsic thermal ripples of free-standing graphene, which contributes to this strain sensitivity. Our results demonstrate that the wettability of 2D membranes is governed not only by their chemistry but also by their dynamic morphology, introducing a new class of wetting behaviour unique to atomically thin materials that offers an additional explanation for variability in experimental measurements. These findings reveal that mechanical strain may be a practical route to controlling wetting in 2D nanomaterials-based technologies, with promising consequences for nanofluidic and nano-filtration applications.},

keywords = {Machine Learning Potentials, Two-dimensional Materials, Water, Water at Interfaces},

pubstate = {published},

tppubtype = {article}

}
David Mendive-Tapia, Christoph Schran, Banshi Das, Fabien Gatti, Markus Schröder, Dominik Marx, Oriol Vendrell
Deciphering the infrared spectrum of the hydrated proton using full-dimensional quantum dynamics Journal Article
In: Nat. Chem., 2026.
Abstract | Links | BibTeX | Tags: Computational Chemistry, Quantum Dynamics, Water
@article{MendiveTapia2026/10.1038/s41557-026-02209-3,

title = {Deciphering the infrared spectrum of the hydrated proton using full-dimensional quantum dynamics},

author = {David Mendive-Tapia and Christoph Schran and Banshi Das and Fabien Gatti and Markus Schröder and Dominik Marx and Oriol Vendrell},

url = {https://www.nature.com/articles/s41557-026-02209-3},

doi = {10.1038/s41557-026-02209-3},

year  = {2026},

date = {2026-07-27},

journal = {Nat. Chem.},

abstract = {The nature of the aqueous proton has been traditionally interpreted through two limiting structural motifs: the Zundel and Eigen cations. However, experimental infrared (IR) spectra of the solvated proton reveal a far more dynamic character, as evidenced by distinct intensity modulations within the characteristic continuum absorption band. In fact, recent ultrafast two-dimensional IR spectroscopy suggests that solvation-induced structural distortions around H2O⋯H+⋯OH2 motifs critically shape the IR response. Here we investigate the role of such asymmetry through full-dimensional quantum dynamics simulations of the extended Zundel complex H⁺(H₂O)₆, which structurally encompasses both Zundel and Eigen motifs. Systematic removal of one water molecule from the second solvation shell gradually introduces deviations from the perfectly symmetric Zundel-like complex towards Eigen-like spectral features. These results provide a direct map between the asymmetric solvation environment and the structural response of the first and second solvation shells of the aqueous proton, offering a structural and dynamical basis for understanding how this asymmetry governs proton mobility in aqueous environments.},

keywords = {Computational Chemistry, Quantum Dynamics, Water},

pubstate = {published},

tppubtype = {article}

}

Xavier R. Advincula, Yair Litman, Kara D. Fong, William C. Witt, Christoph Schran, Angelos Michaelides
How reactive is water at the nanoscale and how to control it? Journal Article
In: Sci. Adv., vol. 12, iss. 26, pp. eaeb5772, 2026.
Abstract | Links | BibTeX | Tags: Chemical Reactivity, Confinement, Machine Learning Potentials, Two-dimensional Materials, Water, Water at Interfaces
@article{Advincula2026/10.1126/sciadv.aeb5772,

title = {How reactive is water at the nanoscale and how to control it?},

author = {Xavier R. Advincula and Yair Litman and Kara D. Fong and William C. Witt and Christoph Schran and Angelos Michaelides},

url = {https://www.science.org/doi/10.1126/sciadv.aeb5772},

doi = {10.1126/sciadv.aeb5772},

year  = {2026},

date = {2026-06-24},

journal = {Sci. Adv.},

volume = {12},

issue = {26},

pages = {eaeb5772},

abstract = {Nanoconfined water plays a key role in nanofluidics, electrochemistry, and catalysis, yet its reactivity remains a matter of debate. Prior studies have reported both enhanced and suppressed water self-dissociation relative to the bulk, but without a consistent explanation. Here, using enhanced sampling molecular dynamics with machine-learned potentials trained at first-principles accuracy, we investigate dissociation behavior in water confined within two-dimensional slit pores and nanodroplets, using graphene and hexagonal boron nitride as model materials. We find that reactivity is extremely sensitive to water density, geometry, and surface chemistry, among other factors. Despite this complexity, we show that chemical potential, together with interfacial interactions, governs dissociation trends and explains the variability observed in prior studies. Within this framework, when confined water is compared to the bulk at equivalent chemical potential, corresponding to thermodynamic equilibrium with a bulk reservoir, its reactivity remains essentially unchanged; rather, differences arise when the systems are compared at different chemical potentials or under distinct interfacial conditions. This thermodynamic perspective reconciles previous contradictions and reveals how nanoscale environments can drastically shift water reactivity. Our findings provide molecular-level insight and offer a design lever for modulating water chemistry at the nanoscale.},

keywords = {Chemical Reactivity, Confinement, Machine Learning Potentials, Two-dimensional Materials, Water, Water at Interfaces},

pubstate = {published},

tppubtype = {article}

}

Samuel W. Coles, Amir Hajibabaei, Venkat Kapil, Xavier R. Advincula, Christoph Schran, Stephen J. Cox, Angelos Michaelides
Nanoconfined superionic water is a molecular superionic Journal Article
In: Sci. Adv., vol. 12, no. 15, pp. eadz6392, 2026.
Abstract | Links | BibTeX | Tags: Charge Transfer, Confinement, Hydrogen Bonding, Water
@article{Coles2026/10.1126/sciadv.adz6392,

title = {Nanoconfined superionic water is a molecular superionic},

author = {Samuel W. Coles and Amir Hajibabaei and Venkat Kapil and Xavier R. Advincula and Christoph Schran and Stephen J. Cox and Angelos Michaelides},

url = {https://www.science.org/doi/full/10.1126/sciadv.adz6392},

doi = {10.1126/sciadv.adz6392},

year  = {2026},

date = {2026-04-10},

journal = {Sci. Adv.},

volume = {12},

number = {15},

pages = {eadz6392},

abstract = {Superionic ice, where water molecules dissociate into a lattice of oxygen ions and a rapidly diffusing "gas" of protons, represents a state of matter with broad implications for planetary interiors and energy applications. Recently, a nanoconfined superionic state of water has been predicted which, in contrast, is composed of intact water molecules. Here, we apply machine learning and electronic structure simulations to establish how nanoconfined water can be both molecular and superionic, providing more general insights into superionic behavior. Similar to bulk ice and other superionic materials, nanoconfined water conducts via concerted chain-like proton migrations, which cause the rapid propagation of defects. However, unlike other molecular phases of water, its exceptional conductivity arises from the activation of the Grotthuss mechanism by (i) low barriers to proton transfer and (ii) a flexible hydrogen bonded network. We propose that these are two key characteristics of fast ionic conduction in molecular superionics.},

keywords = {Charge Transfer, Confinement, Hydrogen Bonding, Water},

pubstate = {published},

tppubtype = {article}

}

Yixuan Feng, Xavier R. Advincula, Hongwei Fang, Christoph Schran
Clay edges are dynamic proton-conducting networks modulated by structure and pH Journal Article
In: J. Phys. Chem. Lett., vol. 17, iss. 9, pp. 2679–2688, 2026.
Abstract | Links | BibTeX | Tags: Charge Transfer, Chemical Reactivity, Ions in Water, Water, Water at Interfaces
@article{Feng2026/10.1021/acs.jpclett.5c03748,

title = {Clay edges are dynamic proton-conducting networks modulated by structure and pH},

author = {Yixuan Feng and Xavier R. Advincula and Hongwei Fang and Christoph Schran},

url = {https://pubs.acs.org/doi/full/10.1021/acs.jpclett.5c03748},

doi = {10.1021/acs.jpclett.5c03748},

year  = {2026},

date = {2026-02-22},

urldate = {2026-02-22},

journal = {J. Phys. Chem. Lett.},

volume = {17},

issue = {9},

pages = {2679–2688},

abstract = {Montmorillonite, a ubiquitous clay mineral, plays a vital role in geochemical and environmental processes due to its chemically complex edge surfaces. However, the molecular-scale acid-base reactivity of these interfaces remains poorly understood due to the limitations of both experimental resolution and conventional simulations. Here, we employ machine learning potentials with first-principles accuracy to perform nanosecond-scale molecular dynamics simulations of montmorillonite nanoparticles across a range of pH. Our results reveal clear amphoteric behavior, with edge sites undergoing protonation in acidic environments and deprotonation in basic conditions, accompanied by pH-dependent surface charge regulation. Even at neutral pH, spontaneous and directional proton transfer events are common, proceeding via both direct and solvent-mediated pathways. These findings demonstrate that montmorillonite edges are not static arrays of hydroxyl groups but dynamic, proton-conducting networks whose reactivity and charge state are modulated by local structure and solution conditions. This work offers a molecular-level framework for understanding proton transport and buffering in clay-water systems, with broad implications for catalysis, ion exchange, and environmental remediation.},

keywords = {Charge Transfer, Chemical Reactivity, Ions in Water, Water, Water at Interfaces},

pubstate = {published},

tppubtype = {article}

}

Xavier R. Advincula, Christoph Schran, Angelos Michaelides
When is nanoconfined water different from interfacial water? Journal Article
In: Faraday Discuss., 2026.
Abstract | Links | BibTeX | Tags: Confinement, Hydrogen Bonding, Two-dimensional Materials, Water, Water at Interfaces
@article{Advincula2026/10.1039/d5fd00165j,

title = {When is nanoconfined water different from interfacial water?},

author = {Xavier R. Advincula and Christoph Schran and Angelos Michaelides},

url = {https://pubs.rsc.org/en/content/articlelanding/2026/fd/d5fd00165j},

doi = {10.1039/D5FD00165J},

year  = {2026},

date = {2026-01-14},

urldate = {2026-01-14},

journal = {Faraday Discuss.},

abstract = {Water behaves very differently at surfaces and under extreme confinement, but the boundary between these two regimes has remained unclear. Despite evidence that interfacial effects persist under sub-nanometre confinement, the molecular-scale behaviour and its evolution with slit width remain unclear. Here, we use machine-learning molecular dynamics with first-principles accuracy to probe water at graphene surfaces across slit widths ranging from the open-interface limit to angstrom-scale confinement. We find that water undergoes a sharp structural transition: when three or more water layers fit between the walls, the structure of the graphene-water interface is effectively indistinguishable from that in an open system, with density layering, hydrogen bonding, and orientational ordering retaining interfacial character. Below this threshold, however, angstrom-scale confinement strongly reorganises the liquid, producing enhanced ordering, a restructured hydrogen-bond network, and modified orientational motifs. These results establish a molecular-level picture that clearly separates interfacial behaviour from genuine nanoconfinement and provide guidance for predicting and controlling the structure of water in nanoscale solid-liquid environments.},

keywords = {Confinement, Hydrogen Bonding, Two-dimensional Materials, Water, Water at Interfaces},

pubstate = {published},

tppubtype = {article}

}
2025

Niamh O’Neill, Benjamin X. Shi, William J. Baldwin, William C. Witt, Gábor Csányi, Julian D. Gale, Angelos Michaelides, Christoph Schran
Towards routine condensed phase simulations with delta-learned coupled cluster accuracy: Application to liquid water Journal Article
In: J. Chem. Theory Comput., 2025.
Abstract | Links | BibTeX | Tags: Coupled Cluster (CC), Machine Learning Potentials, Water
@article{ONeill2025/10.1021/acs.jctc.5c01377,

title = {Towards routine condensed phase simulations with delta-learned coupled cluster accuracy: Application to liquid water},

author = {Niamh O’Neill and Benjamin X. Shi and William J. Baldwin and William C. Witt and Gábor Csányi and Julian D. Gale and Angelos Michaelides and Christoph Schran},

url = {https://pubs.acs.org/doi/full/10.1021/acs.jctc.5c01377},

doi = {10.1021/acs.jctc.5c01377},

year  = {2025},

date = {2025-11-07},

journal = {J. Chem. Theory Comput.},

abstract = {Simulating liquid water to an accuracy that matches its wealth of available experimental data requires both precise electronic structure methods and reliable sampling of nuclear (quantum) motion. This is challenging because applying the electronic structure method of choice, coupled cluster theory with single, double, and perturbative triple excitations [CCSD(T)] to condensed phase systems, is currently limited by its computational cost and complexity. Recent tour de force efforts have demonstrated that this accuracy can indeed bring simulated liquid water into close agreement with experiment using machine learning potentials (MLPs). However, achieving this remains far from routine, requiring large datasets and significant computational cost. In this work, we introduce a practical approach that combines developments in MLPs with local correlation approximations to enable routine CCSD(T)-level simulations of liquid water. When combined with nuclear quantum effects, we achieve agreement with experiments for structural and transport properties. Importantly, the approach also handles constant-pressure simulations, enabling MLP-based CCSD(T) models to predict isothermal–isobaric bulk properties, such as water’s density maximum, in close agreement with experiment. Encompassing tests across electronic structure, datasets, and MLP architecture, this work provides a practical blueprint towards routinely developing CCSD(T)-based MLPs for the condensed phase.},

keywords = {Coupled Cluster (CC), Machine Learning Potentials, Water},

pubstate = {published},

tppubtype = {article}

}
Xavier R. Advincula, Kara D. Fong, Angelos Michaelides, Christoph Schran
Protons accumulate at the graphene–water interface Journal Article
In: ACS Nano, vol. 19, iss. 18, pp. 17728–17737, 2025.
Abstract | Links | BibTeX | Tags: Hydrogen Bonding, Machine Learning Potentials, Water, Water at Interfaces
@article{Advincula2025/10.1021/acsnano.5c02053,

title = {Protons accumulate at the graphene–water interface},

author = {Xavier R. Advincula and Kara D. Fong and Angelos Michaelides and Christoph Schran},

url = {https://pubs.acs.org/doi/full/10.1021/acsnano.5c02053},

doi = {10.1021/acsnano.5c02053},

year  = {2025},

date = {2025-04-28},

urldate = {2025-04-28},

journal = {ACS Nano},

volume = {19},

issue = {18},

pages = {17728–17737},

abstract = {Water’s ability to autoionize into hydroxide and hydronium ions profoundly influences surface properties, rendering interfaces either basic or acidic. While it is well-established that protons show an affinity to the air–water interface, a critical knowledge gap exists in technologically relevant surfaces like the graphene–water interface. Here we use machine learning-based simulations with first-principles accuracy to unravel the behavior of hydroxide and hydronium ions at the graphene–water interface. Our findings reveal that protons accumulate at the graphene–water interface, with the hydronium ion predominantly residing in the first contact layer of water. In contrast, the hydroxide ion exhibits a bimodal distribution, found both near the surface and further away from it. Analysis of the underlying electronic structure reveals local polarization effects, resulting in counterintuitive charge rearrangement. Proton propensity to the graphene–water interface challenges the interpretation of surface experiments and is expected to have far-reaching consequences for ion conductivity, interfacial reactivity, and proton-mediated processes.},

keywords = {Hydrogen Bonding, Machine Learning Potentials, Water, Water at Interfaces},

pubstate = {published},

tppubtype = {article}

}
2024

Niamh O'Neill, Christoph Schran, Stephen J. Cox, Angelos Michaelides
Crumbling crystals: on the dissolution mechanism of NaCl in water Journal Article
In: Phys. Chem. Chem. Phys., vol. 26, iss. 42, pp. 26933-26942, 2024.
Abstract | Links | BibTeX | Tags: Ions in Water, Machine Learning Potentials, Water
@article{ONeill2024/10.1039/d4cp03115f,

title = {Crumbling crystals: on the dissolution mechanism of NaCl in water},

author = {Niamh O'Neill and Christoph Schran and Stephen J. Cox and Angelos Michaelides},

url = {https://pubs.rsc.org/en/content/articlelanding/2024/cp/d4cp03115f},

doi = {10.1039/d4cp03115f},

year  = {2024},

date = {2024-10-30},

urldate = {2024-10-30},

journal = {Phys. Chem. Chem. Phys.},

volume = {26},

issue = {42},

pages = {26933-26942},

abstract = {Dissolution of ionic salts in water is ubiquitous, particularly for NaCl. However, an atomistic scale understanding of the process remains elusive. Simulations lend themselves conveniently to studying dissolution since they provide the spatio-temporal resolution that can be difficult to obtain experimentally. Nevertheless, the complexity of various inter- and intra-molecular interactions require careful treatment and long time scale simulations, both of which are typically hindered by computational expense. Here, we use advances in machine learning potential methodology to resolve at an ab initio level of theory the dissolution mechanism of NaCl in water. The picture that emerges is that of a steady ion-wise unwrapping of the crystal preceding its rapid disintegration, reminiscent of crumbling. The onset of crumbling can be explained by a strong increase in the ratio of the surface area to volume of the crystal. Overall, dissolution comprises a series of highly dynamical microscopic sub-processes, resulting in an inherently stochastic mechanism. These atomistic level insights contribute to the general understanding of dissolution mechanisms in other crystals, and the methodology is primed for more complex systems of recent interest such as water/salt interfaces under flow and salt crystals under confinement.},

keywords = {Ions in Water, Machine Learning Potentials, Water},

pubstate = {published},

tppubtype = {article}

}
Samuel G.H. Brookes, Venkat Kapil, Christoph Schran, Angelos Michaelides
The wetting of H₂O by CO₂ Journal Article
In: J. Chem. Phys., vol. 161, no. 8, pp. 084711, 2024, ISSN: 10897690.
Abstract | Links | BibTeX | Tags: Machine Learning Potentials, Water, Water at Interfaces
@article{Brookes2024/10.1063/5.0224230,

title = {The wetting of H₂O by CO₂},

author = {Samuel G.H. Brookes and Venkat Kapil and Christoph Schran and Angelos Michaelides},

url = {/aip/jcp/article/161/8/084711/3309975/The-wetting-of-H2O-by-CO2},

doi = {10.1063/5.0224230},

issn = {10897690},

year  = {2024},

date = {2024-08-28},

urldate = {2024-08-28},

journal = {J. Chem. Phys.},

volume = {161},

number = {8},

pages = {084711},

publisher = {American Institute of Physics},

abstract = {Biphasic interfaces are complex but fascinating regimes that display a number of properties distinct from those of the bulk. The CO2-H2O interface, in particular, has been the subject of a number of studies on account of its importance for the carbon life cycle as well as carbon capture and sequestration schemes. Despite this attention, there remain a number of open questions on the nature of the CO2-H2O interface, particularly concerning the interfacial tension and phase behavior of CO2 at the interface. In this paper, we seek to address these ambiguities using ab initio-quality simulations. Harnessing the benefits of machine-learned potentials and enhanced statistical sampling methods, we present an ab initio-level description of the CO2-H2O interface. Interfacial tensions are predicted from 1 to 500 bars and found to be in close agreement with experiment at pressures for which experimental data are available. Structural analyses indicate the buildup of an adsorbed, saturated CO2 film forming at a low pressure (20 bars) with properties similar to those of the bulk liquid, but preferential perpendicular alignment with respect to the interface. The CO2 monolayer buildup coincides with a reduced structuring of water molecules close to the interface. This study highlights the predictive nature of machine-learned potentials for complex macroscopic properties of biphasic interfaces, and the mechanistic insight obtained into carbon dioxide aggregation at the water interface is of high relevance for geoscience, climate research, and materials science.},

keywords = {Machine Learning Potentials, Water, Water at Interfaces},

pubstate = {published},

tppubtype = {article}

}
Pavan Ravindra, Xavier R. Advincula, Christoph Schran, Angelos Michaelides, Venkat Kapil
Quasi-one-dimensional hydrogen bonding in nanoconfined ice Journal Article
In: Nat. Commun., vol. 15, no. 1, pp. 1–9, 2024, ISSN: 20411723.
Abstract | Links | BibTeX | Tags: Confinement, Hydrogen Bonding, Machine Learning Potentials, Water
@article{Ravindra2024/10.1038/s41467-024-51124-z,

title = {Quasi-one-dimensional hydrogen bonding in nanoconfined ice},

author = {Pavan Ravindra and Xavier R. Advincula and Christoph Schran and Angelos Michaelides and Venkat Kapil},

url = {https://www.nature.com/articles/s41467-024-51124-z},

doi = {10.1038/s41467-024-51124-z},

issn = {20411723},

year  = {2024},

date = {2024-08-24},

urldate = {2024-08-24},

journal = {Nat. Commun.},

volume = {15},

number = {1},

pages = {1–9},

publisher = {Nature Publishing Group},

abstract = {The Bernal-Fowler ice rules stipulate that each water molecule in an ice crystal should form four hydrogen bonds. However, in extreme or constrained conditions, the arrangement of water molecules deviates from conventional ice rules, resulting in properties significantly different from bulk water. In this study, we employ machine learning-driven first-principles simulations to identify a new stabilization mechanism in nanoconfined ice phases. Instead of forming four hydrogen bonds, nanoconfined crystalline ice can form a quasi-one-dimensional hydrogen-bonded structure that exhibits only two hydrogen bonds per water molecule. These structures consist of strongly hydrogen-bonded linear chains of water molecules that zig-zag along one dimension, stabilized by van der Waals interactions that stack these chains along the other dimension. The unusual interplay of hydrogen bonding and van der Waals interactions in nanoconfined ice results in atypical proton behavior such as potential ferroelectric behavior, low dielectric response, and long-range proton dynamics.},

keywords = {Confinement, Hydrogen Bonding, Machine Learning Potentials, Water},

pubstate = {published},

tppubtype = {article}

}
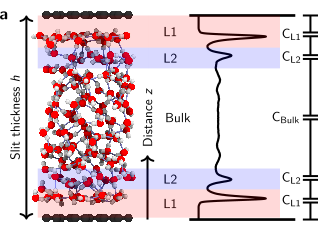
Thomas Dufils, Christoph Schran, Ji Chen, Andre K. Geim, Laura Fumagalli, Angelos Michaelides
Origin of dielectric polarization suppression in confined water from first principles Journal Article
In: Chem. Sci., vol. 15, iss. 2, pp. 516–527, 2024.
Abstract | Links | BibTeX | Tags: Ab-Initio Molecular Dynamics (AIMD), Confinement, Hydrogen Bonding, Water
@article{Dufils2024/10.1039/D3SC04740G,

title = {Origin of dielectric polarization suppression in confined water from first principles},

author = {Thomas Dufils and Christoph Schran and Ji Chen and Andre K. Geim and Laura Fumagalli and Angelos Michaelides},

url = {http://dx.doi.org/10.1039/D3SC04740G},

doi = {10.1039/D3SC04740G},

year  = {2024},

date = {2024-01-01},

urldate = {2024-01-01},

journal = {Chem. Sci.},

volume = {15},

issue = {2},

pages = {516–527},

publisher = {The Royal Society of Chemistry},

abstract = {It has long been known that the dielectric constant of confined water should be different from that in bulk. Recent experiments have shown that it is vanishingly small, however the origin of the phenomenon remains unclear. Here we used ab initio molecular dynamics simulations (AIMD) and AIMD-trained machine-learning potentials to understand water's structure and electronic properties underpinning this effect. For the graphene and hexagonal boron-nitride substrates considered, we find that it originates in the spontaneous anti-parallel alignment of the water dipoles in the first two water layers near the solid interface. The interfacial layers exhibit net ferroelectric ordering, resulting in an overall anti-ferroelectric arrangement of confined water. Together with constrained hydrogen-bonding orientations, this leads to much reduced out-of-plane polarization. Furthermore, we directly contrast AIMD and simple classical force-field simulations, revealing important differences. This work offers insight into a property of water that is critical in modulating surface forces, the electric-double-layer formation and molecular solvation, and shows a way to compute it.},

keywords = {Ab-Initio Molecular Dynamics (AIMD), Confinement, Hydrogen Bonding, Water},

pubstate = {published},

tppubtype = {article}

}
2023
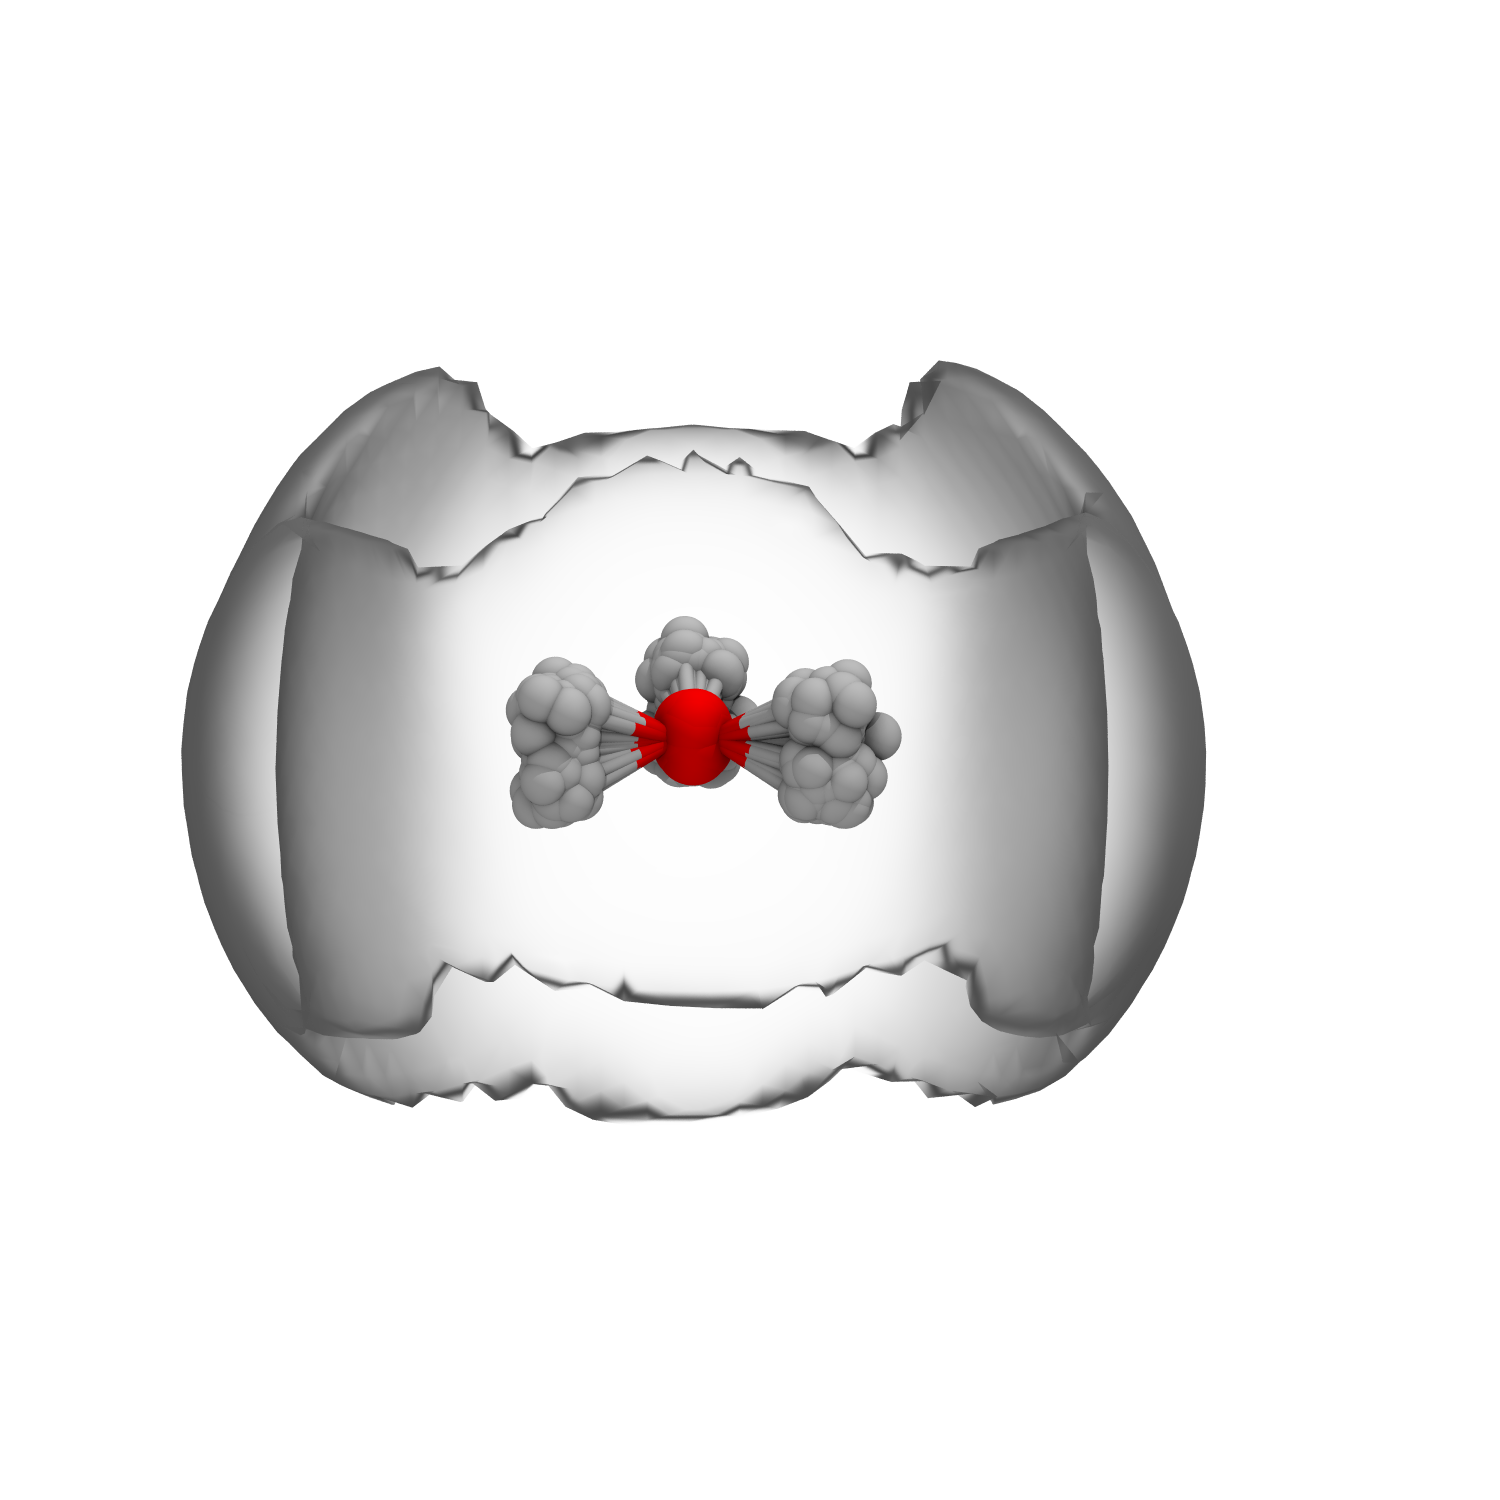
Julia A. Davies, Christoph Schran, Fabien Brieuc, Dominik Marx, Andrew M. Ellis
Onset of rotational decoupling for a molecular ion solvated in helium: From tags to rings and shells Journal Article
In: Phys. Rev. Lett., vol. 130, iss. 8, pp. 083001, 2023.
Links | BibTeX | Tags: Nuclear Quantum Effects (NQEs), Path Integral Molecular Dynamics (PIMD), Superfluidity, Water
@article{Schran2023/10.1103/PhysRevLett.130.083001,

title = {Onset of rotational decoupling for a molecular ion solvated in helium: From tags to rings and shells},

author = {Julia A. Davies and Christoph Schran and Fabien Brieuc and Dominik Marx and Andrew M. Ellis},

doi = {10.1103/PhysRevLett.130.083001},

year  = {2023},

date = {2023-02-01},

urldate = {2023-02-01},

journal = {Phys. Rev. Lett.},

volume = {130},

issue = {8},

pages = {083001},

publisher = {American Physical Society},

keywords = {Nuclear Quantum Effects (NQEs), Path Integral Molecular Dynamics (PIMD), Superfluidity, Water},

pubstate = {published},

tppubtype = {article}

}
2022
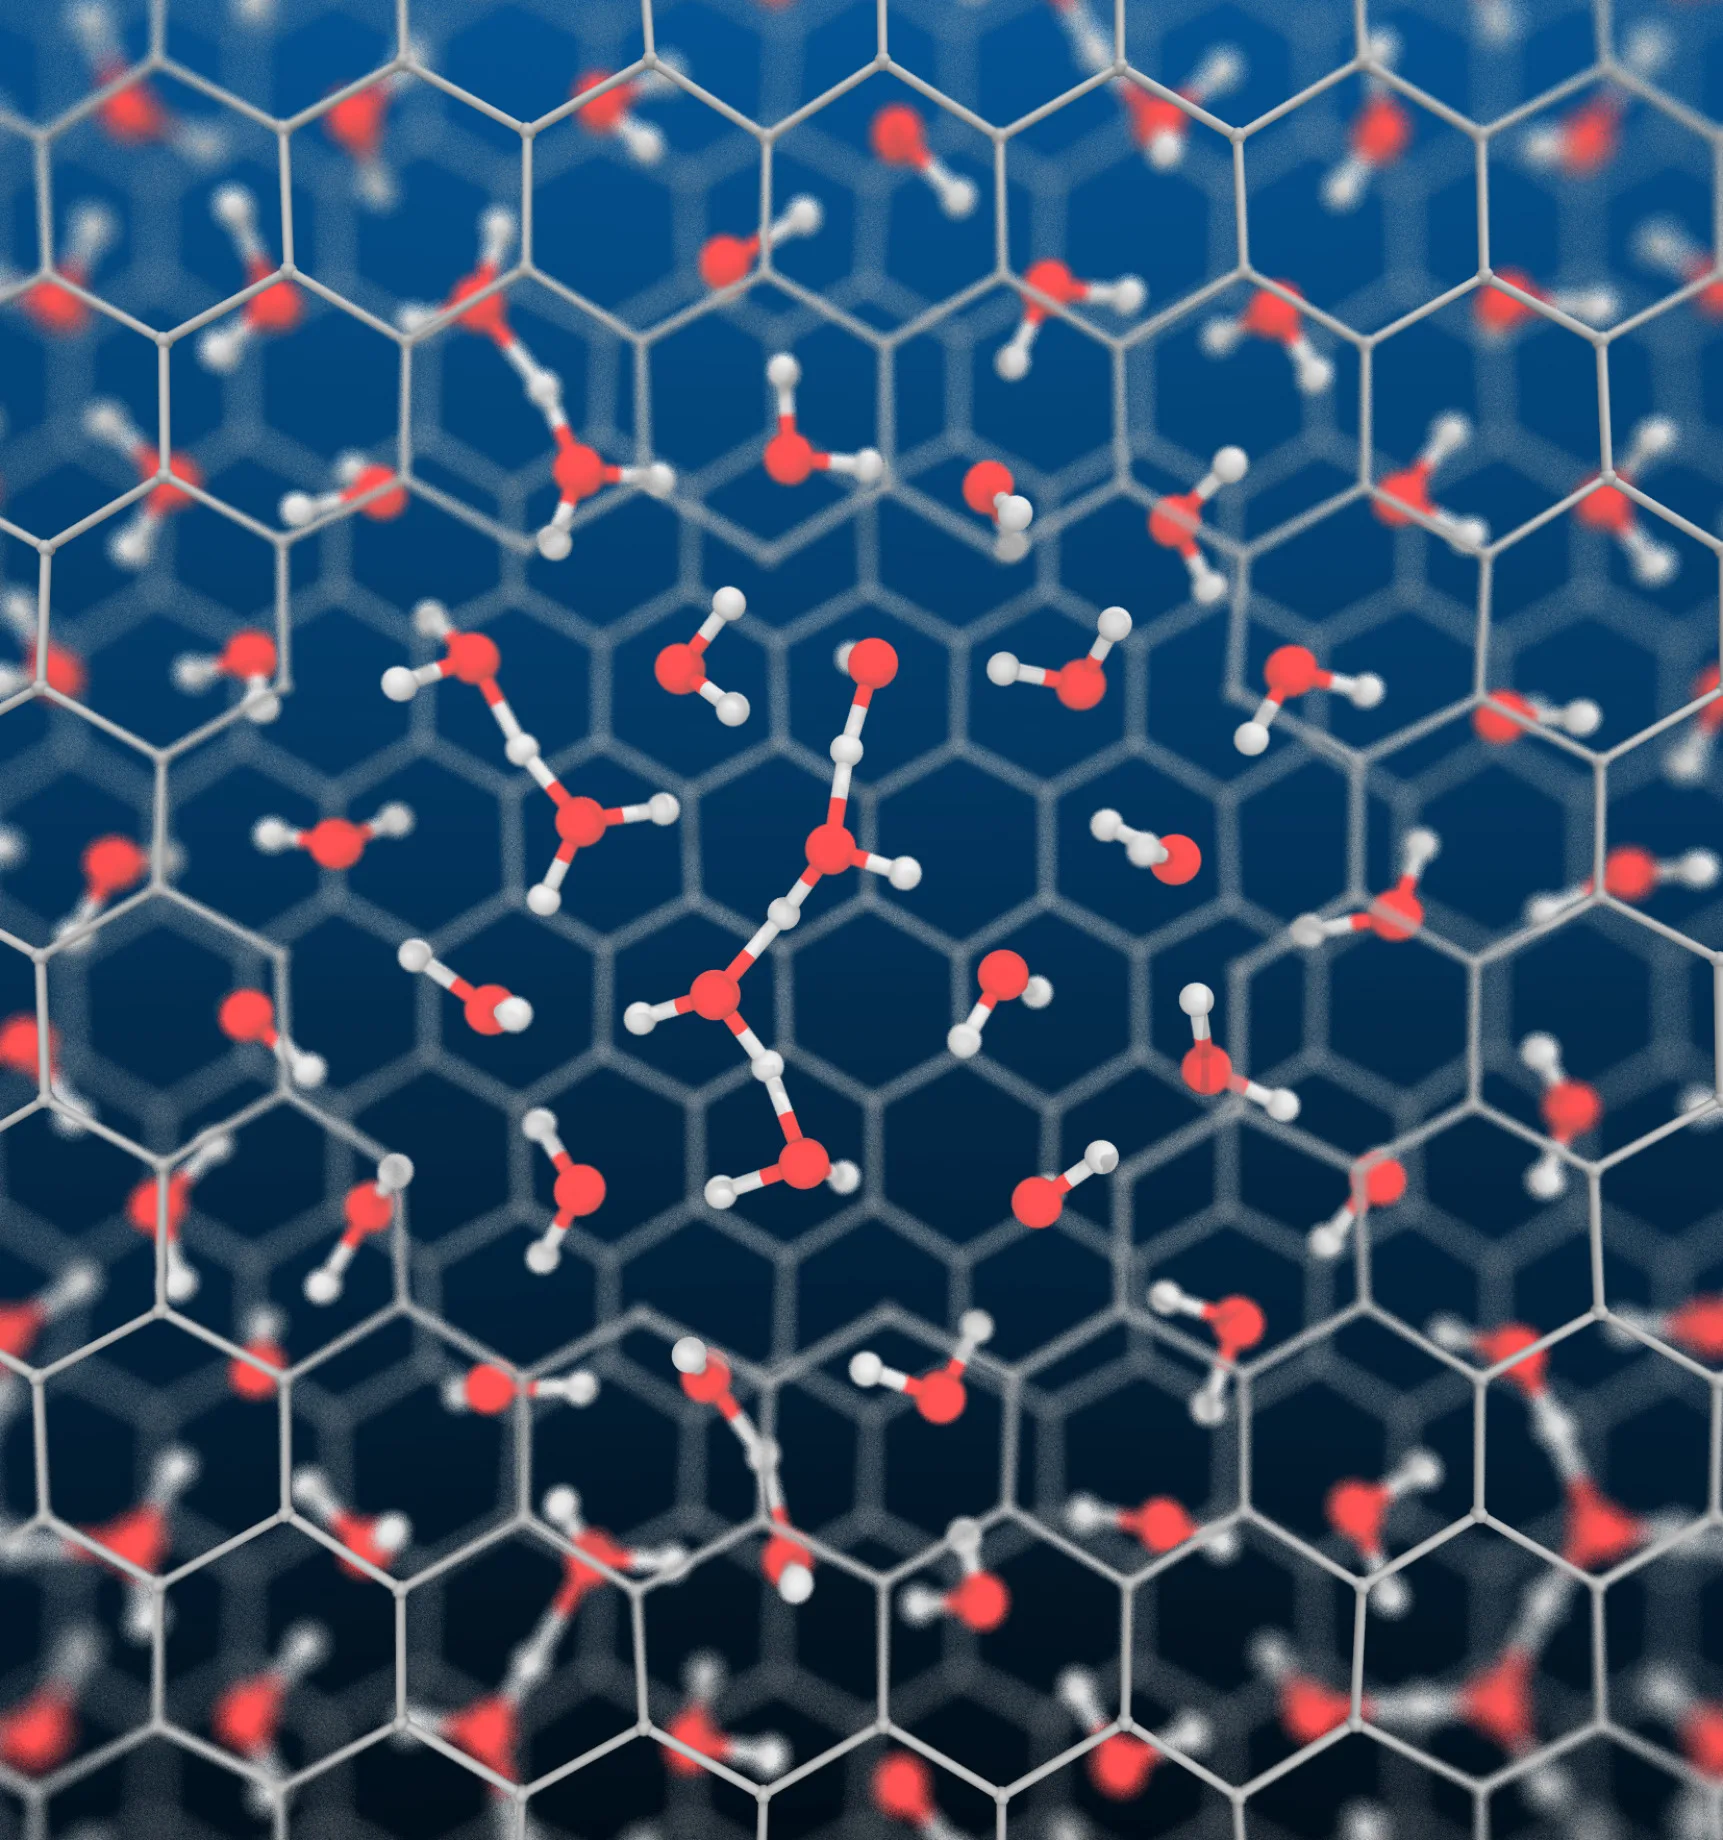
Venkat Kapil, Christoph Schran, Andrea Zen, Ji Chen, Chris J. Pickard, Angelos Michaelides
The first-principles phase diagram of monolayer nanoconfined water Journal Article
In: Nature, vol. 609, pp. 512-516, 2022, ISSN: 1476-4687.
Abstract | Links | BibTeX | Tags: Confinement, Machine Learning Potentials, Water
@article{Kapil2022/10.1038/S41586-022-05036-X,

title = {The first-principles phase diagram of monolayer nanoconfined water},

author = {Venkat Kapil and Christoph Schran and Andrea Zen and Ji Chen and Chris J. Pickard and Angelos Michaelides},

doi = {10.1038/S41586-022-05036-X},

issn = {1476-4687},

year  = {2022},

date = {2022-09-01},

urldate = {2022-09-01},

journal = {Nature},

volume = {609},

pages = {512-516},

publisher = {Nature Publishing Group},

abstract = {Water in nanoscale cavities is ubiquitous and of central importance to everyday phenomena in geology and biology. However, the properties of nanoscale water can be substantially different from those of bulk water, as shown, for example, by the anomalously low dielectric constant of water in nanochannels1, near frictionless water flow2 or the possible existence of a square ice phase3. Such properties suggest that nanoconfined water could be engineered for technological applications in nanofluidics4, electrolyte materials5 and water desalination6. Unfortunately, challenges in experimentally characterizing water at the nanoscale and the high cost of first-principles simulations have prevented the molecular-level understanding required to control the behaviour of water. Here we combine a range of computational approaches to enable a first-principles-level investigation of a single layer of water within a graphene-like channel. We find that monolayer water exhibits surprisingly rich and diverse phase behaviour that is highly sensitive to temperature and the van der Waals pressure acting within the nanochannel. In addition to multiple molecular phases with melting temperatures varying non-monotonically by more than 400 kelvins with pressure, we predict a hexatic phase, which is an intermediate between a solid and a liquid, and a superionic phase with a high electrical conductivity exceeding that of battery materials. Notably, this suggests that nanoconfinement could be a promising route towards superionic behaviour under easily accessible conditions. Monolayer water exhibits rich and diverse phase behaviour that is highly sensitive to temperature and the van der Waals pressure acting within the nanochannel.},

keywords = {Confinement, Machine Learning Potentials, Water},

pubstate = {published},

tppubtype = {article}

}
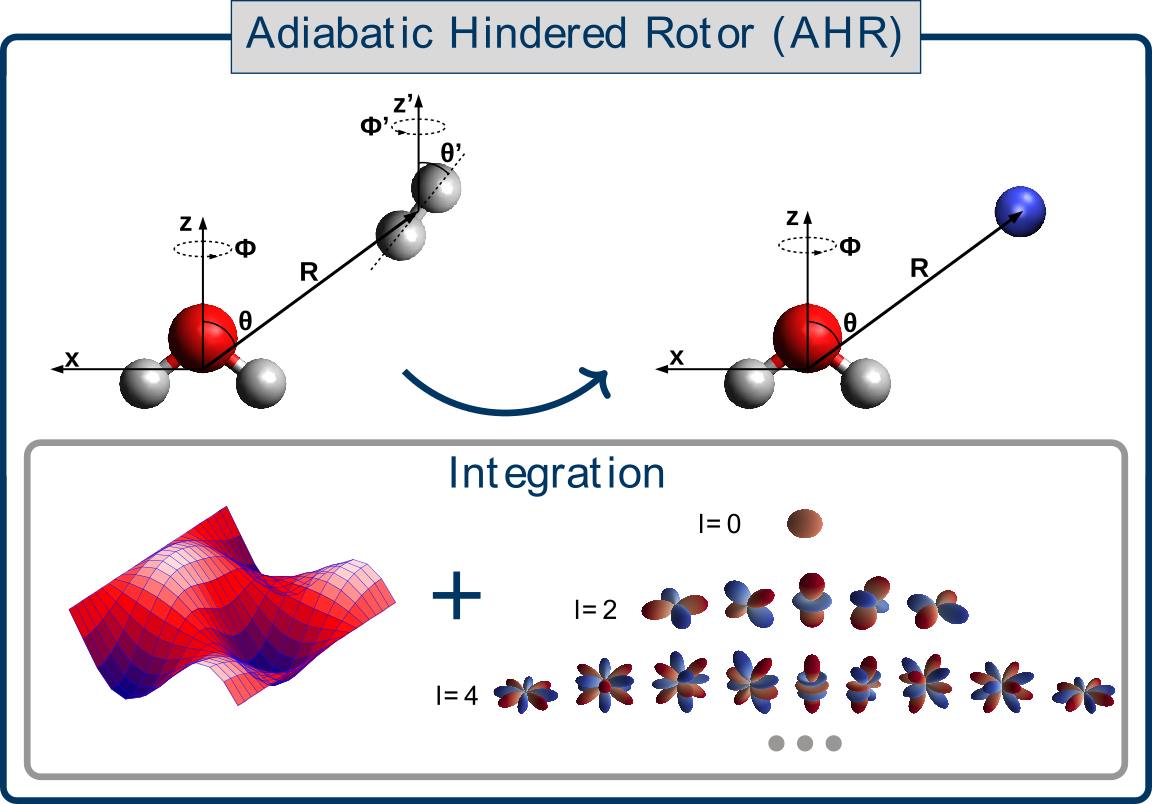
Laura Durán Caballero, Christoph Schran, Fabien Brieuc, Dominik Marx
Neural network interaction potentials for para-hydrogen with flexible molecules Journal Article
In: J. Chem. Phys., vol. 157, no. 7, pp. 074302, 2022, ISSN: 0021-9606.
Abstract | Links | BibTeX | Tags: Machine Learning Potentials, Nuclear Quantum Effects (NQEs), Superfluidity, Water
@article{Duran2022/10.1063/5.0100953,

title = {Neural network interaction potentials for para-hydrogen with flexible molecules},

author = {Laura Durán Caballero and Christoph Schran and Fabien Brieuc and Dominik Marx},

doi = {10.1063/5.0100953},

issn = {0021-9606},

year  = {2022},

date = {2022-08-01},

urldate = {2022-08-01},

journal = {J. Chem. Phys.},

volume = {157},

number = {7},

pages = {074302},

publisher = {AIP Publishing LLCAIP Publishing},

abstract = {The study of molecular impurities in para-hydrogen (pH2) clusters is key to push forward our understanding of intra- and intermolecular interactions including their impact on the superfluid response of this bosonic quantum solvent. This includes tagging with one or very few pH2, the microsolvation regime, and matrix isolation. However, the fundamental coupling between the bosonic pH2 environment and the (ro-)vibrational motion of molecular impurities remains poorly understood. Quantum simulations can in provide the necessary atomistic insight, but very accurate descriptions of the involved interactions are required. Here, we present a data-driven approach for the generation of impurity-pH2 interaction potentials based on machine learning techniques which retain the full flexibility of the impurity. We employ the well-established adiabatic hindered rotor (AHR) averaging technique to include the impact of the nuclear spin statistics on the symmetry-allowed rotational quantum numbers of pH2. Embedding this averaging procedure within the high-dimensional neural network potential (NNP) framework enables the generation of highly-accurate AHR-averaged NNPs at coupled cluster accuracy, namely CCSD(T*)-F12a/aVTZcp in an automated manner. We apply this methodology to the water and protonated water molecules, as representative cases for quasi-rigid and highly-flexible molecules respectively, and obtain AHR-averaged NNPs that reliably describe the H2O-pH2 and H3O+-pH2 interactions. Using path integral simulations we show for the hydronium cation that umbrella-like tunneling inversion has a strong impact on the first and second pH2 microsolvation shells. The data-driven nature of our protocol opens the door to the study of bosonic pH2 quantum solvation for a wide range of embedded impurities.},

keywords = {Machine Learning Potentials, Nuclear Quantum Effects (NQEs), Superfluidity, Water},

pubstate = {published},

tppubtype = {article}

}
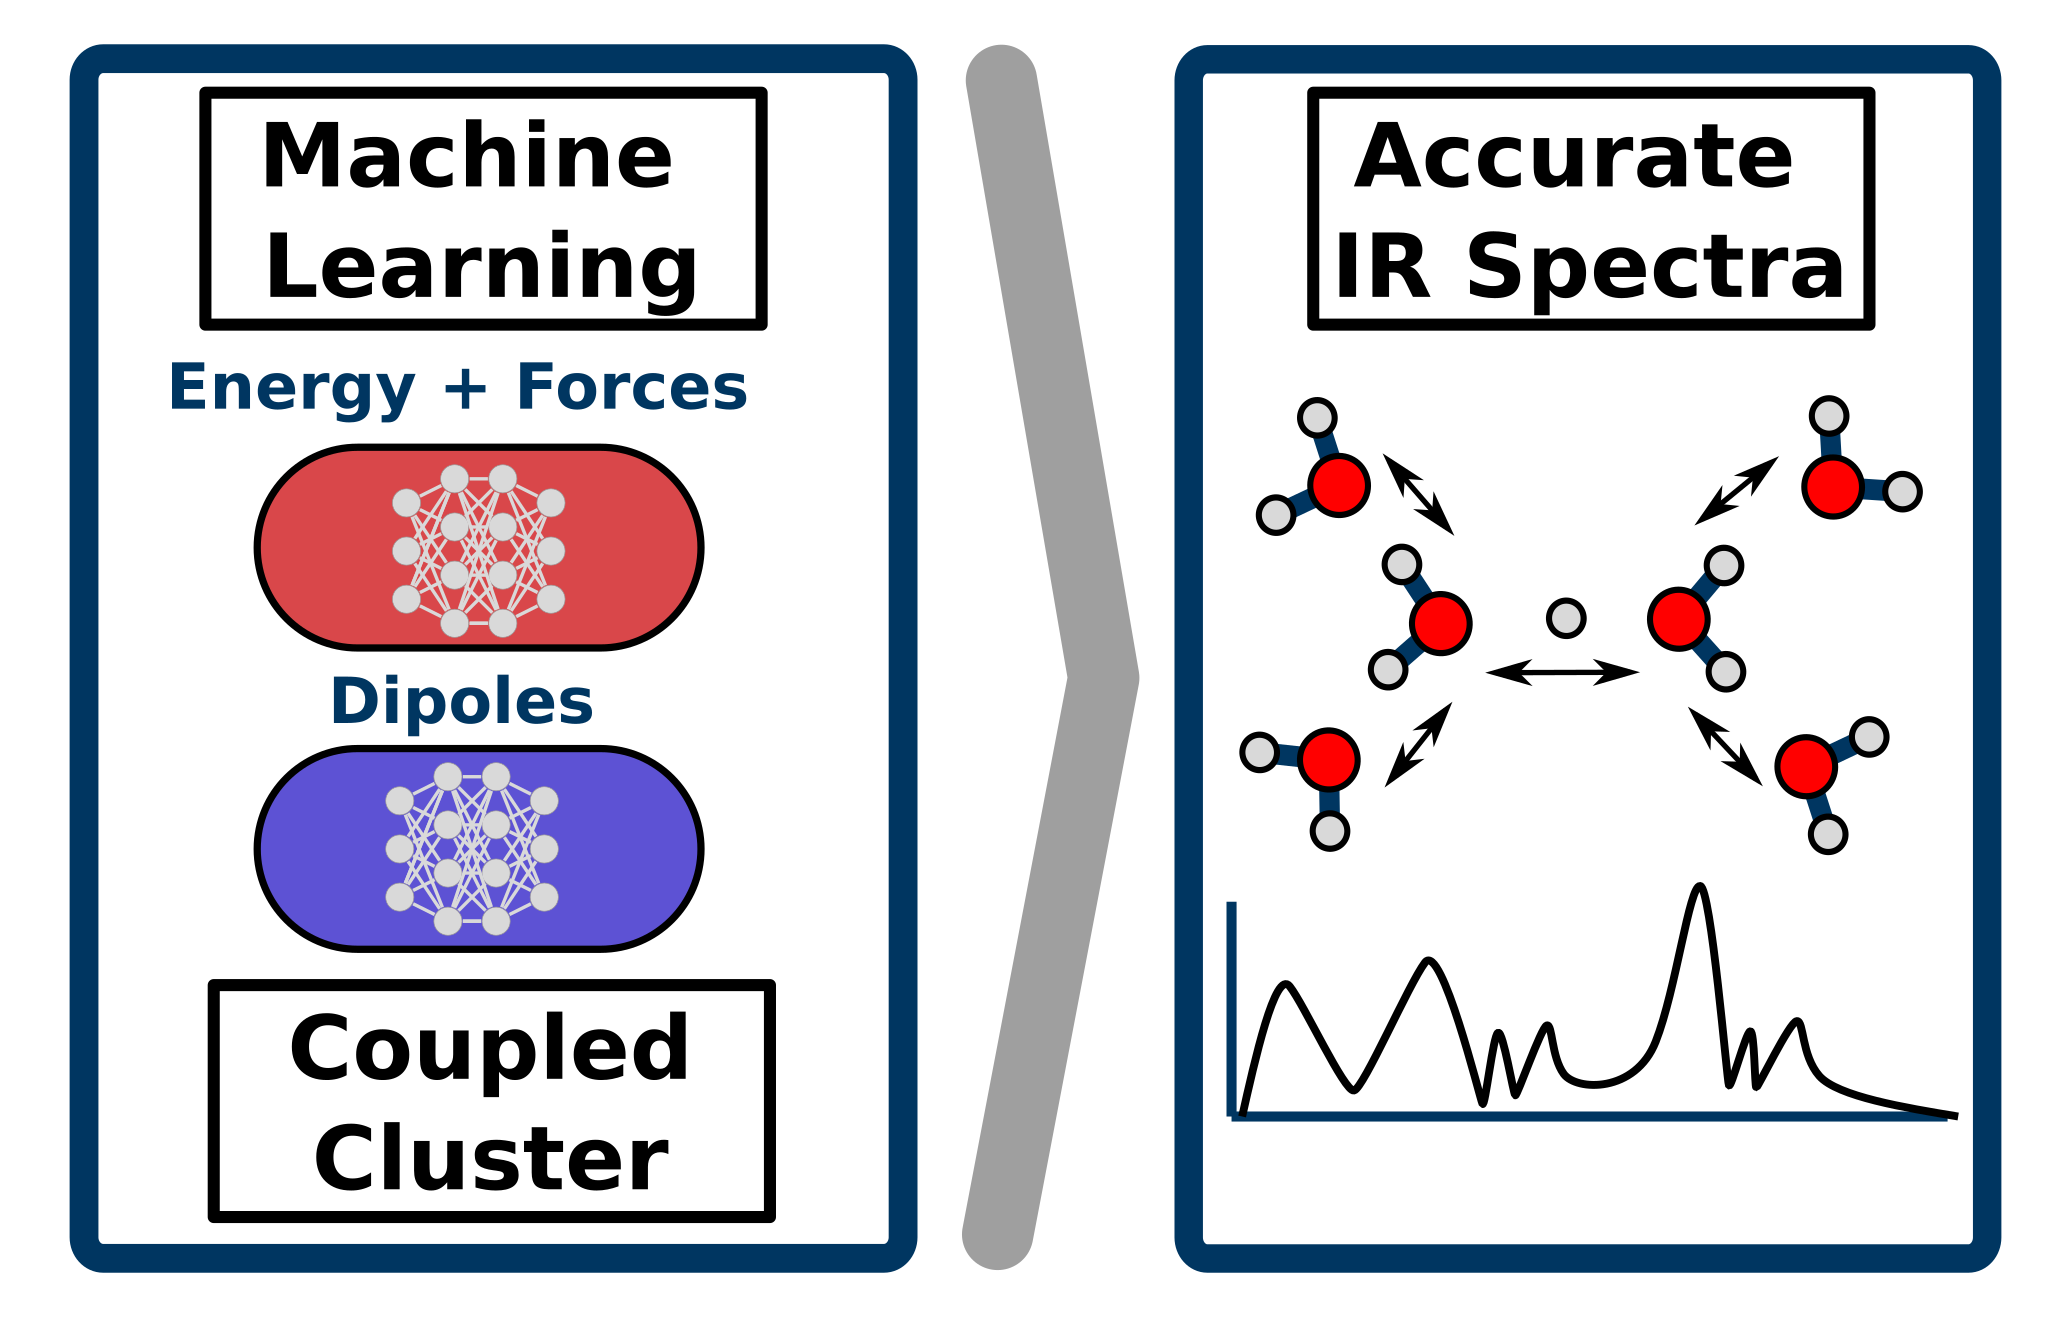
Richard Beckmann, Fabien Brieuc, Christoph Schran, Dominik Marx
Infrared spectra at coupled cluster accuracy from neural network representations Journal Article
In: J. Chem. Theory Comput., 2022, ISSN: 1549-9618.
Abstract | Links | BibTeX | Tags: Coupled Cluster (CC), Machine Learning Potentials, Spectra, Water
@article{Beckmann2022/10.1021/ACS.JCTC.2C00511,

title = {Infrared spectra at coupled cluster accuracy from neural network representations},

author = {Richard Beckmann and Fabien Brieuc and Christoph Schran and Dominik Marx},

doi = {10.1021/ACS.JCTC.2C00511},

issn = {1549-9618},

year  = {2022},

date = {2022-08-01},

urldate = {2022-08-01},

journal = {J. Chem. Theory Comput.},

publisher = {American Chemical Society},

abstract = {Infrared spectroscopy is key to elucidate molecular structures, monitor reactions and observe conformational changes, while providing information on both structural and dynamical properties. This makes the accurate prediction of infrared spectra based on first-principle theories a highly desirable pursuit. Molecular dynamics simulations have proven to be a particularly powerful approach for this task, albeit requiring the computation of energies, forces and dipole moments for a large number of molecular configurations as a function of time. This explains why highly accurate first principles methods, such as coupled cluster theory, have so far been inapplicable for the prediction of fully anharmonic vibrational spectra of large systems at finite temperatures. Here, we push cutting-edge machine learning techniques forward by using neural network representations of energies, forces and in particular dipoles to predict such infrared spectra fully at "gold standard" coupled cluster accuracy as demonstrated for protonated water clusters as large as the protonated water hexamer, in its extended Zundel configuration. Furthermore, we show that this methodology can be used beyond the scope of the data considered during the development of the neural network models, allowing for the computation of finite-temperature infrared spectra of large systems inaccessible to explicit coupled cluster calculations. This substantially expands the hitherto existing limits of accuracy, speed and system size for theoretical spectroscopy and opens up a multitude of avenues for the prediction of vibrational spectra and the understanding of complex intra- and intermolecular couplings.},

keywords = {Coupled Cluster (CC), Machine Learning Potentials, Spectra, Water},

pubstate = {published},

tppubtype = {article}

}
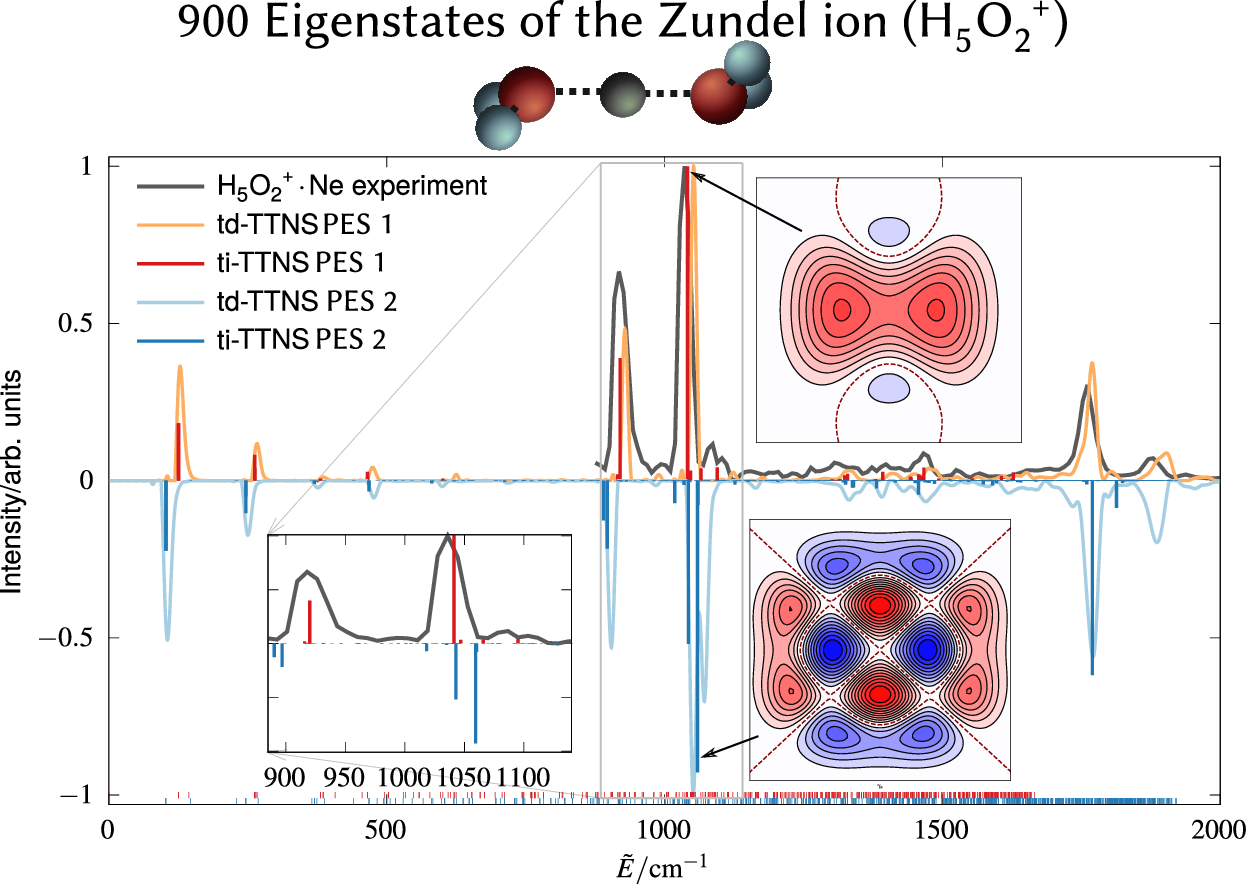
Henrik R Larsson, Markus Schröder, Richard Beckmann, Fabien Brieuc, Christoph Schran, Dominik Marx, Oriol Vendrell
State-resolved infrared spectrum of the protonated water dimer: Revisiting the characteristic proton transfer doublet peak Journal Article
In: Chem. Sci., 2022, ISSN: 2041-6539.
Abstract | Links | BibTeX | Tags: Quantum Dynamics, Spectra, Water
@article{Larsson2022/10.1039/D2SC03189B,

title = {State-resolved infrared spectrum of the protonated water dimer: Revisiting the characteristic proton transfer doublet peak},

author = {Henrik R Larsson and Markus Schröder and Richard Beckmann and Fabien Brieuc and Christoph Schran and Dominik Marx and Oriol Vendrell},

doi = {10.1039/D2SC03189B},

issn = {2041-6539},

year  = {2022},

date = {2022-08-01},

urldate = {2022-08-01},

journal = {Chem. Sci.},

publisher = {The Royal Society of Chemistry},

abstract = {The infrared (IR) spectra of protonated water clusters encode precise information on the dynamics and structure of the hydrated proton. However, the strong anharmonic coupling and quantum effects of these elusive species remain puzzling up to the present day. Here, we report unequivocal evidence that the interplay between the proton transfer and the water wagging motions in the protonated water dimer (Zundel ion) giving rise to the characteristic doublet peak is both more complex and more sensitive to subtle energetic changes than previously thought. In particular, hitherto overlooked low-intensity satellite peaks in the experimental spectrum are now unveiled and mechanistically assigned. Our findings rely on the comparison of IR spectra obtained using two highly accurate potential energy surfaces in conjunction with highly accurate state-resolved quantum simulations. We demonstrate that these high-accuracy simulations are important for providing definite assignments of the complex IR signals of fluxional molecules.},

keywords = {Quantum Dynamics, Spectra, Water},

pubstate = {published},

tppubtype = {article}

}
2021
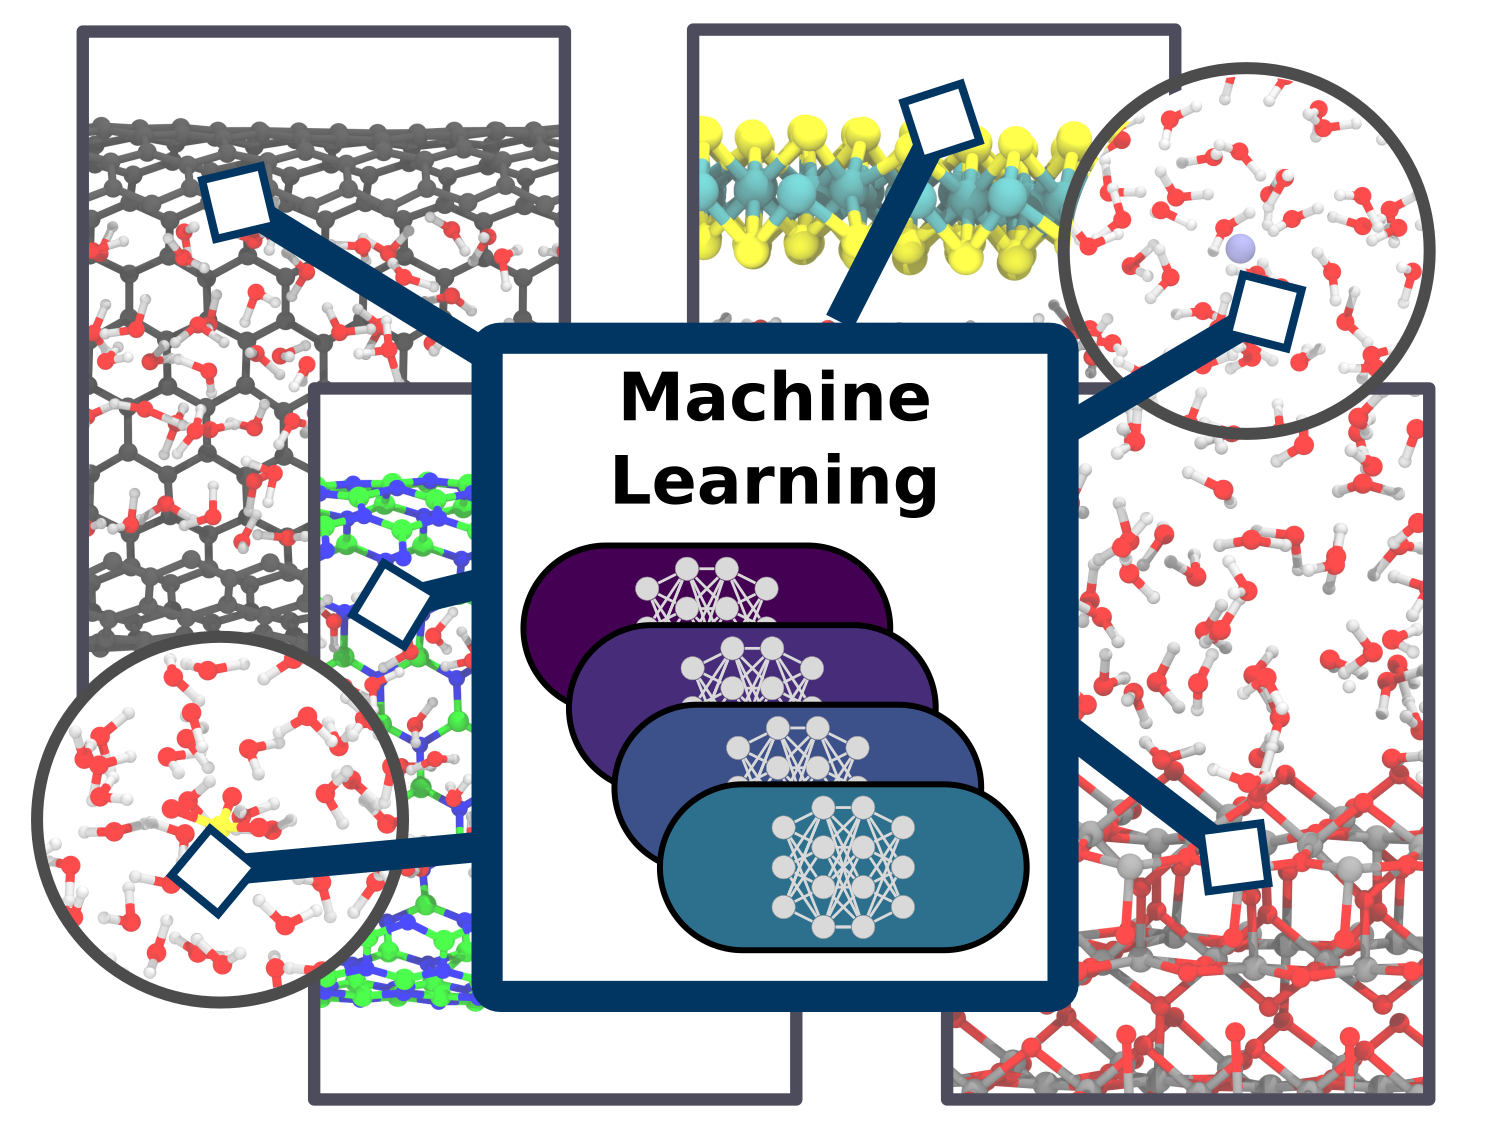
Christoph Schran, Fabian L. Thiemann, Patrick Rowe, Erich A. Müller, Ondrej Marsalek, Angelos Michaelides
Machine learning potentials for complex aqueous systems made simple Journal Article
In: Proc. Natl. Acad. Sci., vol. 118, no. 38, pp. e2110077118, 2021, ISSN: 0027-8424.
Abstract | Links | BibTeX | Tags: Confinement, Ions in Water, Machine Learning Potentials, Water, Water at Interfaces
@article{Schran2021/10.1073/PNAS.2110077118,

title = {Machine learning potentials for complex aqueous systems made simple},

author = {Christoph Schran and Fabian L. Thiemann and Patrick Rowe and Erich A. Müller and Ondrej Marsalek and Angelos Michaelides},

doi = {10.1073/PNAS.2110077118},

issn = {0027-8424},

year  = {2021},

date = {2021-09-01},

urldate = {2021-09-01},

journal = {Proc. Natl. Acad. Sci.},

volume = {118},

number = {38},

pages = {e2110077118},

publisher = {National Academy of Sciences},

abstract = {Simulation techniques based on accurate and efficient representations of potential energy surfaces are urgently needed for the understanding of complex aqueous systems such as solid-liquid interfaces. Here, we present a machine learning framework that enables the efficient development and validation of models for complex aqueous systems. Instead of trying to deliver a globally-optimal machine learning potential, we propose to develop models applicable to specific thermodynamic state points in a simple and user-friendly process. After an initial ab initio simulation, a machine learning potential is constructed with minimum human effort through a data-driven active learning protocol. Such models can afterwards be applied in exhaustive simulations to provide reliable answers for the scientific question at hand. We showcase this methodology on a diverse set of aqueous systems with increasing degrees of complexity. The systems chosen here comprise water with different ions in solution, water on a titanium dioxide surface, as well as water confined in nanotubes and between molybdenum disulfide sheets. Highlighting the accuracy of our approach compared to the ab initio reference, the resulting models are evaluated in detail with an automated validation protocol that includes structural and dynamical properties and the precision of the force prediction of the models. Finally, we demonstrate the capabilities of our approach for the description of water on the rutile titanium dioxide (110) surface to analyze structure and mobility of water on this surface. Such machine learning models provide a straightforward and uncomplicated but accurate extension of simulation time and length scales for complex systems.},

keywords = {Confinement, Ions in Water, Machine Learning Potentials, Water, Water at Interfaces},

pubstate = {published},

tppubtype = {article}

}
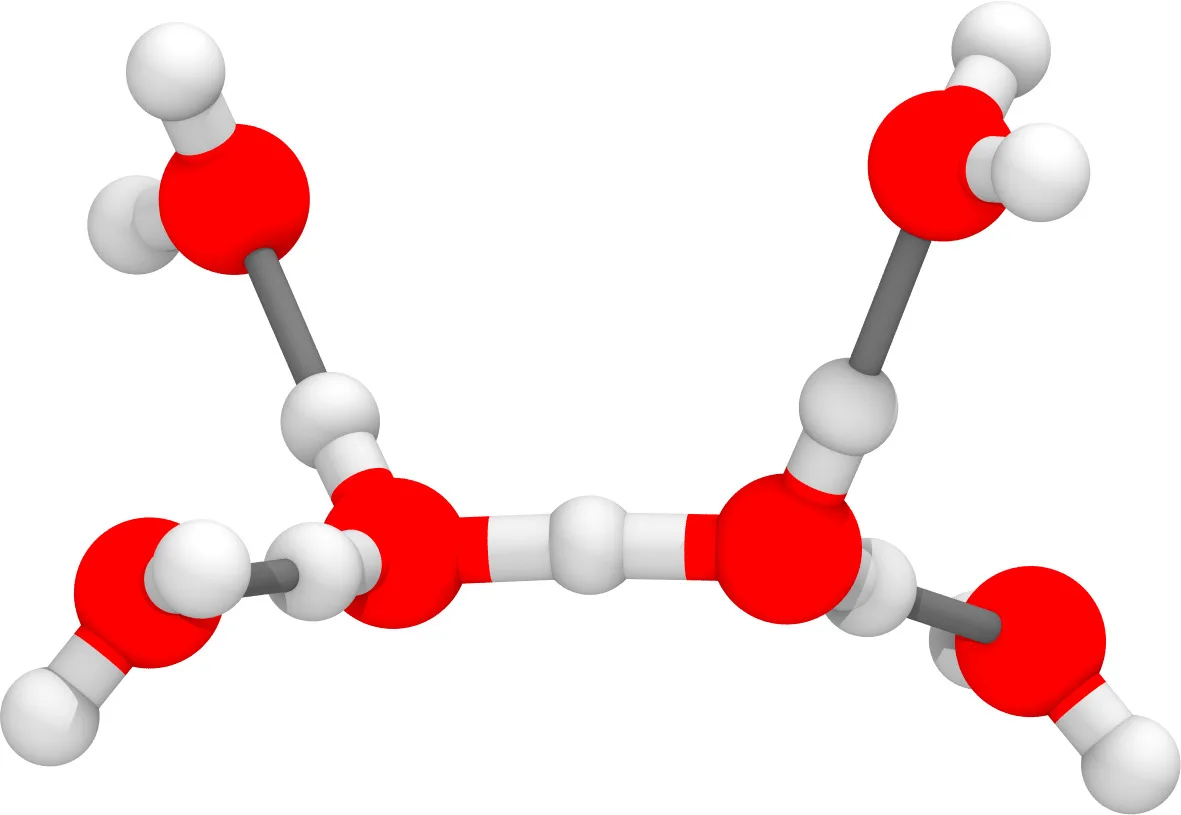
Christoph Schran, Fabien Brieuc, Dominik Marx
Transferability of machine learning potentials: Protonated water neural network potential applied to the protonated water hexamer Journal Article
In: J. Chem. Phys., vol. 154, no. 5, pp. 051101, 2021, ISSN: 10897690.
Abstract | Links | BibTeX | Tags: Coupled Cluster (CC), Hydrogen Bonding, Machine Learning Potentials, Water
@article{Schran2021/10.1063/5.0035438,

title = {Transferability of machine learning potentials: Protonated water neural network potential applied to the protonated water hexamer},

author = {Christoph Schran and Fabien Brieuc and Dominik Marx},

doi = {10.1063/5.0035438},

issn = {10897690},

year  = {2021},

date = {2021-02-01},

urldate = {2021-02-01},

journal = {J. Chem. Phys.},

volume = {154},

number = {5},

pages = {051101},

abstract = {A previously published neural network potential for the description of protonated water clusters up to the protonated water tetramer, H+(H2O)4, at an essentially converged coupled cluster accuracy [C. Schran, J. Behler, and D. Marx, J. Chem. Theory Comput. 16, 88 (2020)] is applied to the protonated water hexamer, H+(H2O)6 - a system that the neural network has never seen before. Although being in the extrapolation regime, it is shown that the potential not only allows for quantum simulations from ultra-low temperatures ∼1 K up to 300 K but is also able to describe the new system very accurately compared to explicit coupled cluster calculations. This transferability of the model is rationalized by the similarity of the atomic environments encountered for the larger cluster compared to the environments in the training set of the model. Compared to the interpolation regime, the quality of the model is reduced by roughly one order of magnitude, but most of the difference to the coupled cluster reference comes from global shifts of the potential energy surface, while local energy fluctuations are well recovered. These results suggest that the application of neural network potentials in extrapolation regimes can provide useful results and might be more general than usually thought.},

keywords = {Coupled Cluster (CC), Hydrogen Bonding, Machine Learning Potentials, Water},

pubstate = {published},

tppubtype = {article}

}
2020
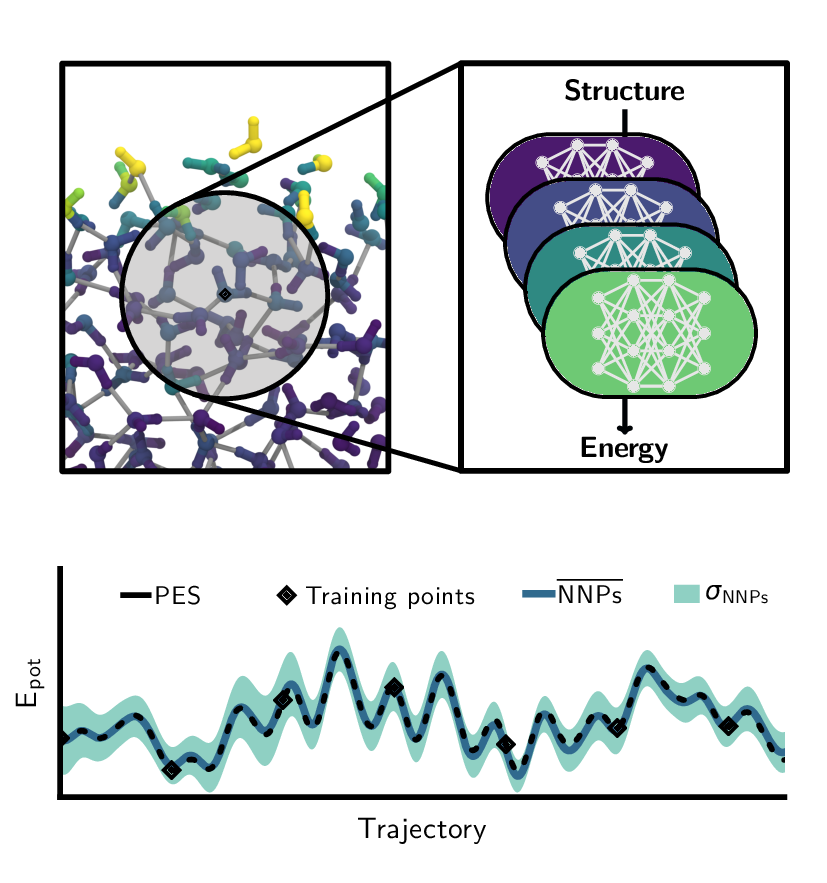
Christoph Schran, Krystof Brezina, Ondrej Marsalek
Committee neural network potentials control generalization errors and enable active learning Journal Article
In: J. Chem. Phys., vol. 153, no. 10, pp. 104105, 2020, ISSN: 10897690.
Abstract | Links | BibTeX | Tags: Machine Learning Potentials, Path Integral Molecular Dynamics (PIMD), Water
@article{Schran2020/10.1063/5.0016004,

title = {Committee neural network potentials control generalization errors and enable active learning},

author = {Christoph Schran and Krystof Brezina and Ondrej Marsalek},

doi = {10.1063/5.0016004},

issn = {10897690},

year  = {2020},

date = {2020-09-01},

urldate = {2020-09-01},

journal = {J. Chem. Phys.},

volume = {153},

number = {10},

pages = {104105},

abstract = {It is well known in the field of machine learning that committee models improve accuracy, provide generalization error estimates, and enable active learning strategies. In this work, we adapt these concepts to interatomic potentials based on artificial neural networks. Instead of a single model, multiple models that share the same atomic environment descriptors yield an average that outperforms its individual members as well as a measure of the generalization error in the form of the committee disagreement. We not only use this disagreement to identify the most relevant configurations to build up the model's training set in an active learning procedure but also monitor and bias it during simulations to control the generalization error. This facilitates the adaptive development of committee neural network potentials and their training sets while keeping the number of ab initio calculations to a minimum. To illustrate the benefits of this methodology, we apply it to the development of a committee model for water in the condensed phase. Starting from a single reference ab initio simulation, we use active learning to expand into new state points and to describe the quantum nature of the nuclei. The final model, trained on 814 reference calculations, yields excellent results under a range of conditions, from liquid water at ambient and elevated temperatures and pressures to different phases of ice, and the air-water interface - all including nuclear quantum effects. This approach to committee models will enable the systematic development of robust machine learning models for a broad range of systems.},

keywords = {Machine Learning Potentials, Path Integral Molecular Dynamics (PIMD), Water},

pubstate = {published},

tppubtype = {article}

}
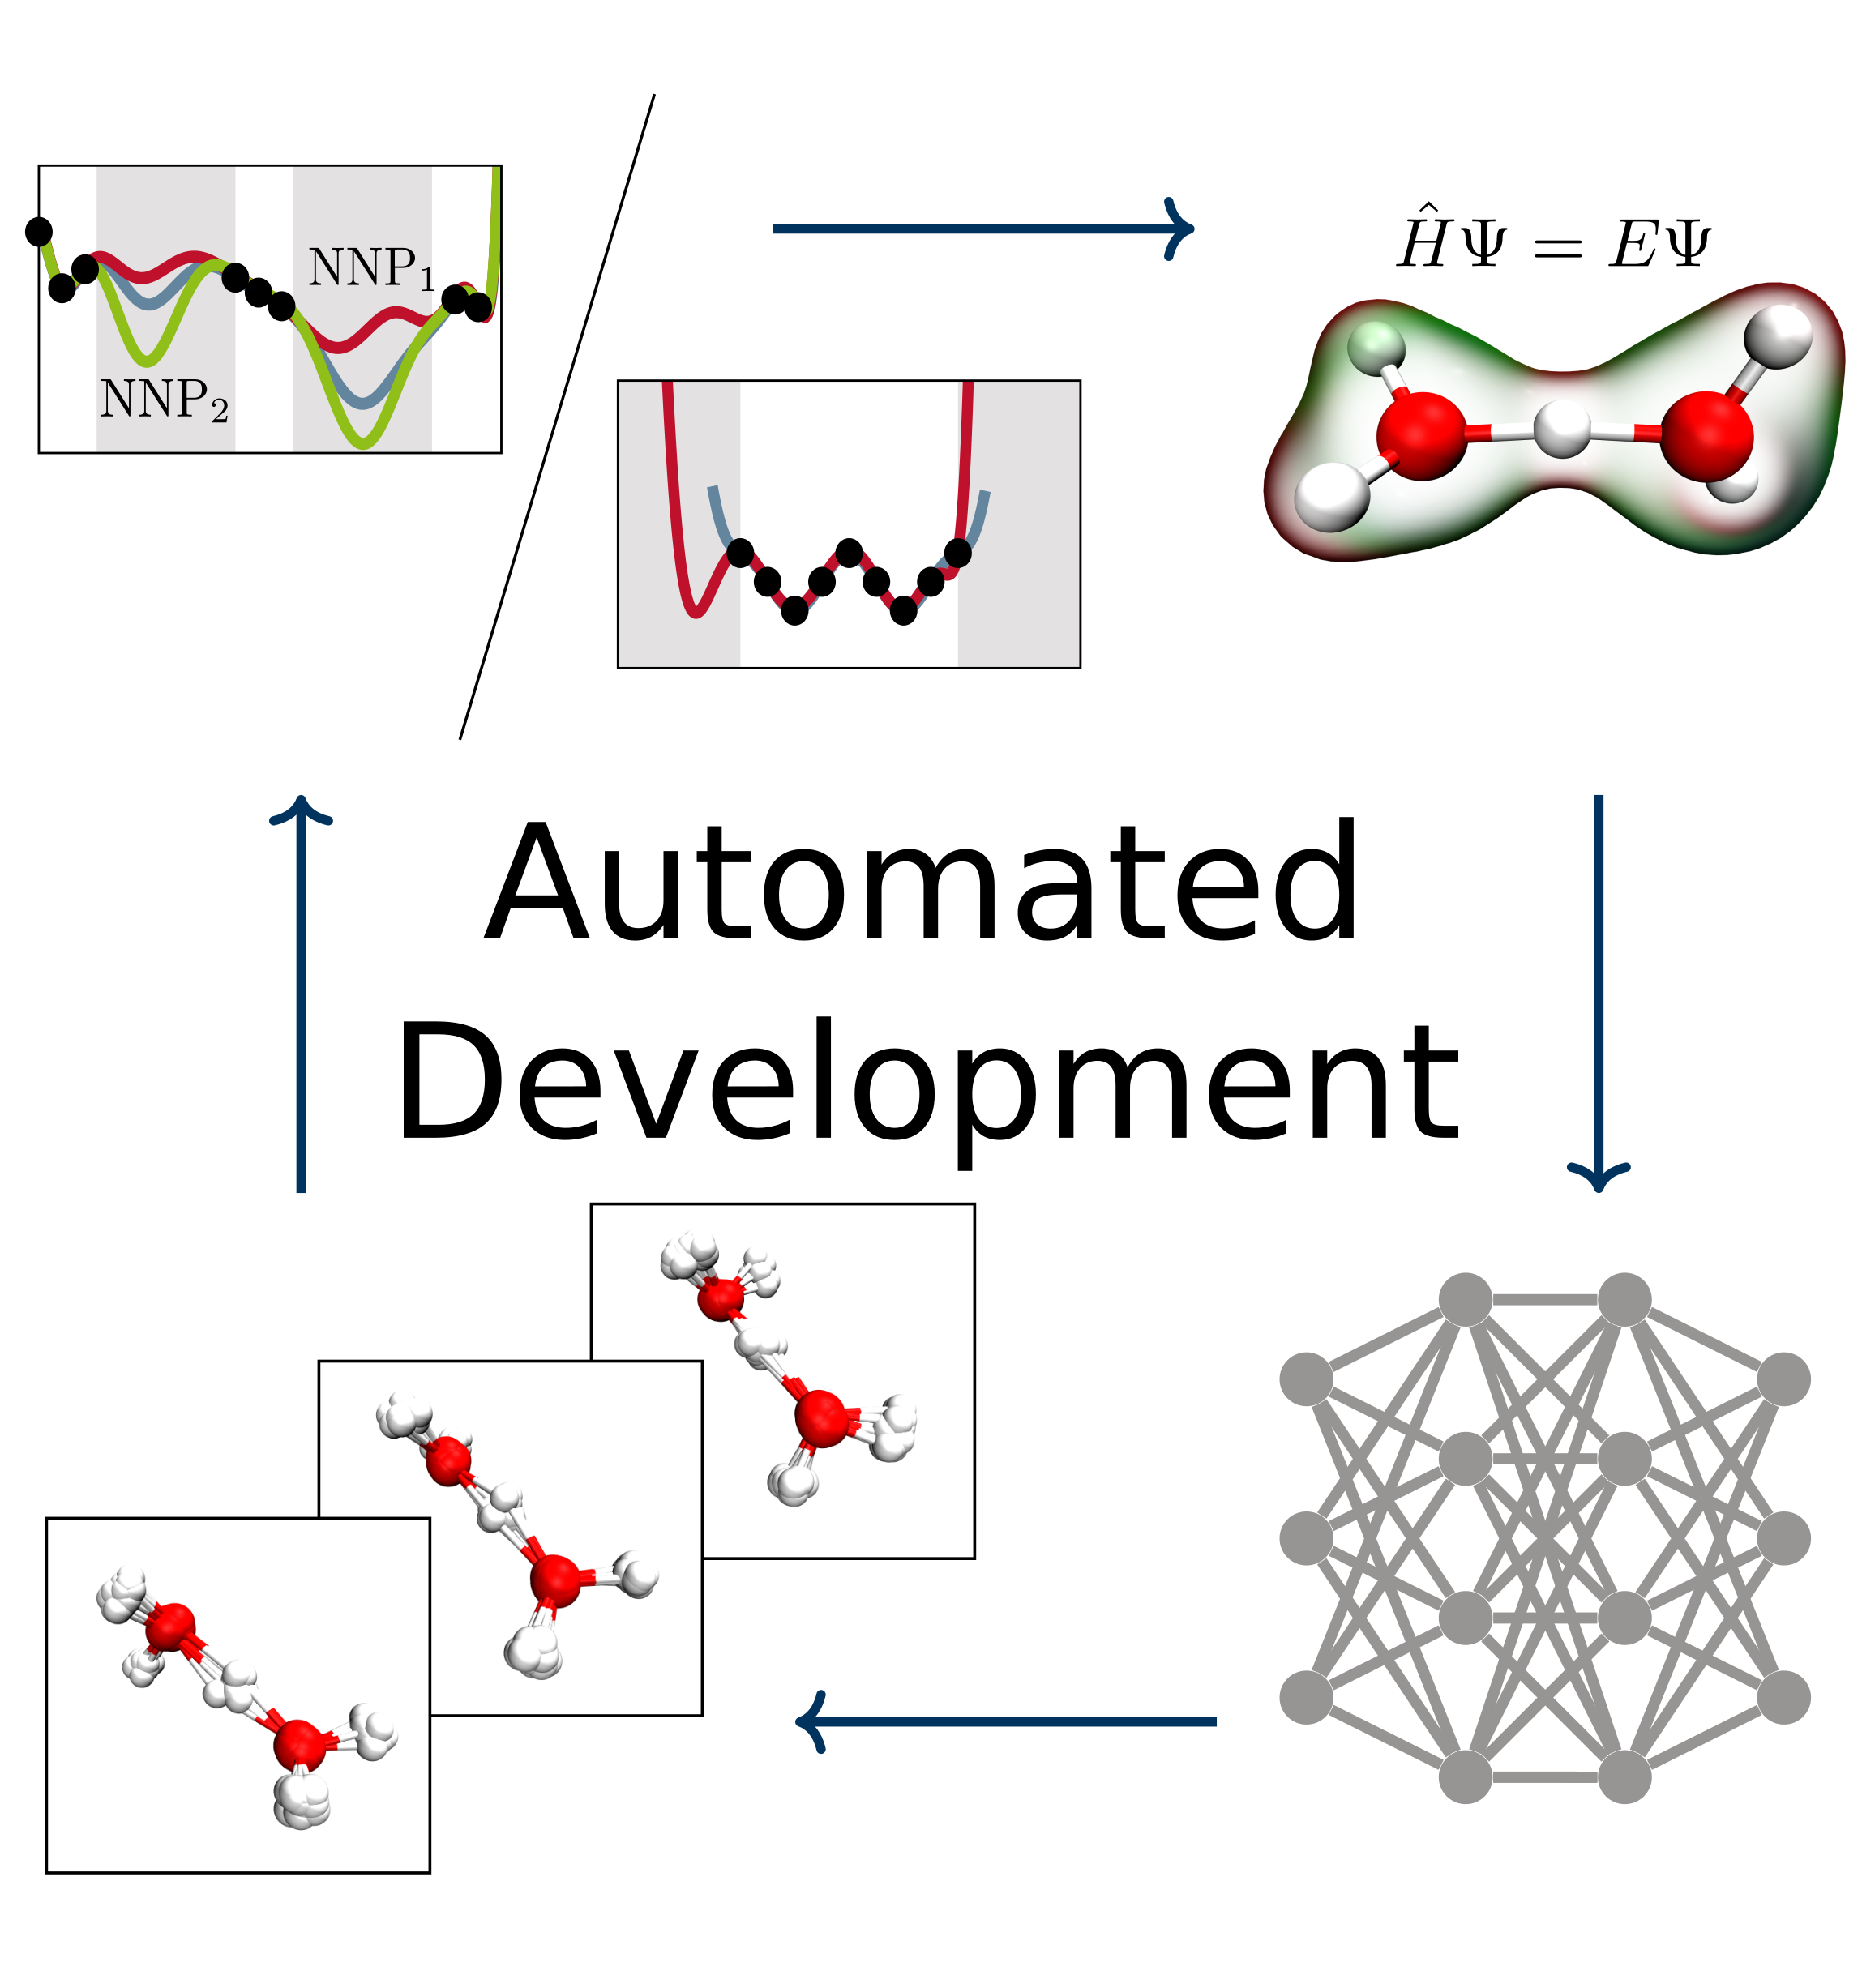
Christoph Schran, Jörg Behler, Dominik Marx
Automated fitting of neural network potentials at coupled cluster accuracy: Protonated water clusters as testing ground Journal Article
In: J. Chem. Theory Comput., vol. 16, no. 1, pp. 88–99, 2020, ISSN: 15499626.
Abstract | Links | BibTeX | Tags: Coupled Cluster (CC), Hydrogen Bonding, Machine Learning Potentials, Water
@article{Schran2020/10.1021/acs.jctc.9b00805,

title = {Automated fitting of neural network potentials at coupled cluster accuracy: Protonated water clusters as testing ground},

author = {Christoph Schran and Jörg Behler and Dominik Marx},

doi = {10.1021/acs.jctc.9b00805},

issn = {15499626},

year  = {2020},

date = {2020-01-01},

urldate = {2020-01-01},

journal = {J. Chem. Theory Comput.},

volume = {16},

number = {1},

pages = {88–99},

abstract = {Highly accurate potential energy surfaces are of key interest for the detailed understanding and predictive modeling of chemical systems. In recent years, several new types of force fields, which are based on machine learning algorithms and fitted to ab initio reference calculations, have been introduced to meet this requirement. Here, we show how high-dimensional neural network potentials can be employed to automatically generate the potential energy surface of finite sized clusters at coupled cluster accuracy, namely CCSD(T*)-F12a/aug-cc-pVTZ. The developed automated procedure utilizes the established intrinsic properties of the model such that the configurations for the training set are selected in an unbiased and efficient way to minimize the computational effort of expensive reference calculations. These ideas are applied to protonated water clusters from the hydronium cation, H3O+, up to the tetramer, H9O4+, and lead to a single potential energy surface that describes all these systems at essentially converged coupled cluster accuracy with a fitting error of 0.06 kJ/mol per atom. The fit is validated in detail for all clusters up to the tetramer and yields reliable results not only for stationary points but also for reaction pathways and intermediate configurations as well as different sampling techniques. Per design, the neural network potentials (NNPs) constructed in this fashion can handle very different conditions including the quantum nature of the nuclei and enhanced sampling techniques covering very low as well as high temperatures. This enables fast and exhaustive exploration of the targeted protonated water clusters with essentially converged interactions. In addition, the automated process will allow one to tackle finite systems much beyond the present case.},

keywords = {Coupled Cluster (CC), Hydrogen Bonding, Machine Learning Potentials, Water},

pubstate = {published},

tppubtype = {article}

}
2019
Christoph Schran, Dominik Marx
Quantum nature of the hydrogen bond from ambient conditions down to ultra-low temperatures Journal Article
In: Phys. Chem. Chem. Phys., vol. 21, no. 45, pp. 24967–24975, 2019, ISSN: 14639076.
Abstract | Links | BibTeX | Tags: Hydrogen Bonding, Nuclear Quantum Effects (NQEs), Path Integral Molecular Dynamics (PIMD), Water
@article{Schran2019/10.1039/C9CP04795F,

title = {Quantum nature of the hydrogen bond from ambient conditions down to ultra-low temperatures},

author = {Christoph Schran and Dominik Marx},

doi = {10.1039/c9cp04795f},

issn = {14639076},

year  = {2019},

date = {2019-10-01},

urldate = {2019-10-01},

journal = {Phys. Chem. Chem. Phys.},

volume = {21},

number = {45},

pages = {24967–24975},

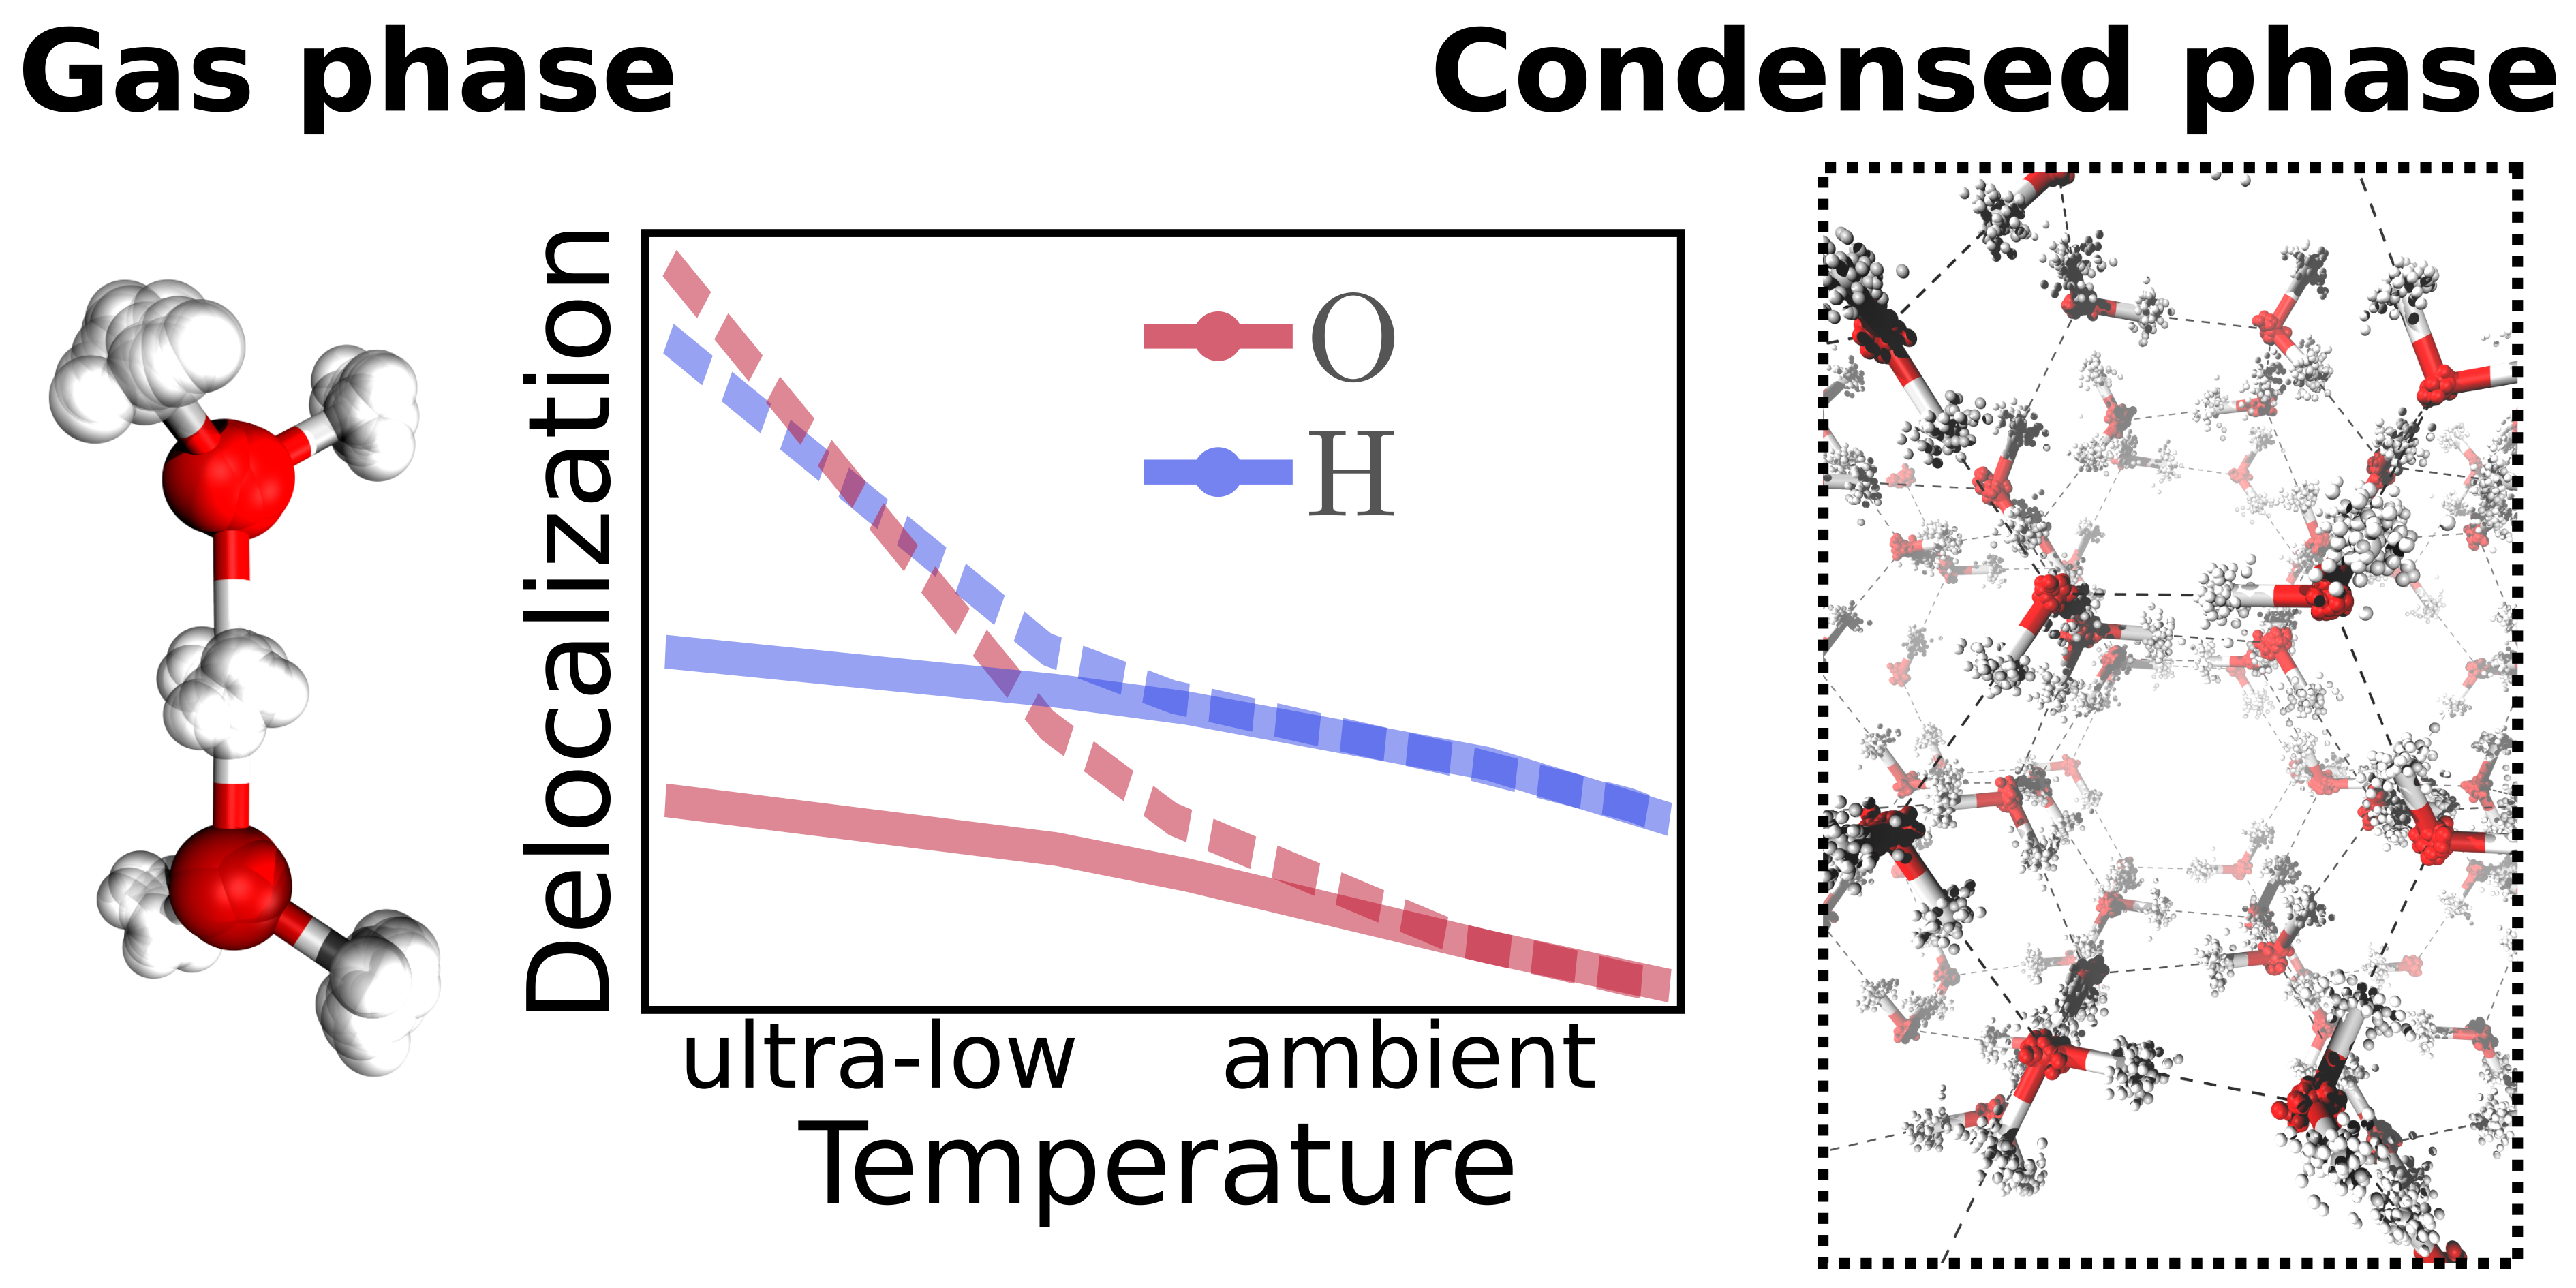
abstract = {Many experimental techniques such as tagging photodissociation and helium nanodroplet isolation spectroscopy operate at very low temperatures in order to investigate hydrogen bonding. To elucidate the differences between such ultra-cold and usual ambient conditions, different hydrogen bonded systems are studied systematically from 300 K down to about 1 K using path integral simulations that explicitly consider both the quantum nature of the nuclei and thermal fluctuations. For this purpose, finite sized water clusters, specifically the water dimer and hexamer, protonated water clusters including the Zundel and Eigen complexes, as well as hexagonal ice as a condensed phase representative are compared directly as a function of temperature. While weaker hydrogen bonds, as present in the neutral systems, show distinct structural differences between ambient conditions and the ultra-cold regime, the stronger hydrogen bonds of the protonated water clusters are less perturbed by temperature compared to their quantum ground state. In all the studied systems, the quantum delocalization of the nuclei is found to vary drastically with temperature. Interestingly, upon reaching temperatures of about 1 K, the spatial quantum delocalization of the heavy oxygens approaches that of the protons for relatively weak spatial constraints, and even significantly exceeds the latter in the case of the centered hydrogen bond in the Zundel complex. These findings are relevant for comparisons between experiments on hydrogen bonding carried out under ultra-cold versus ambient conditions as well as to understand quantum delocalization phenomena of nuclei by seamlessly extending our insights into noncovalent interactions down to ultra-low temperatures.},

keywords = {Hydrogen Bonding, Nuclear Quantum Effects (NQEs), Path Integral Molecular Dynamics (PIMD), Water},

pubstate = {published},

tppubtype = {article}

}
2018
Christoph Schran, Fabien Brieuc, Dominik Marx
Converged colored noise path integral molecular dynamics study of the zundel cation down to ultralow temperatures at coupled cluster accuracy Journal Article
In: J. Chem. Theory Comput., vol. 14, no. 10, pp. 5068–5078, 2018, ISSN: 15499626.
Abstract | Links | BibTeX | Tags: Nuclear Quantum Effects (NQEs), Path Integral Molecular Dynamics (PIMD), Water
@article{Schran2018/10.1021/acs.jctc.8b00705,

title = {Converged colored noise path integral molecular dynamics study of the zundel cation down to ultralow temperatures at coupled cluster accuracy},

author = {Christoph Schran and Fabien Brieuc and Dominik Marx},

doi = {10.1021/acs.jctc.8b00705},

issn = {15499626},

year  = {2018},

date = {2018-09-01},

urldate = {2018-09-01},

journal = {J. Chem. Theory Comput.},

volume = {14},

number = {10},

pages = {5068–5078},

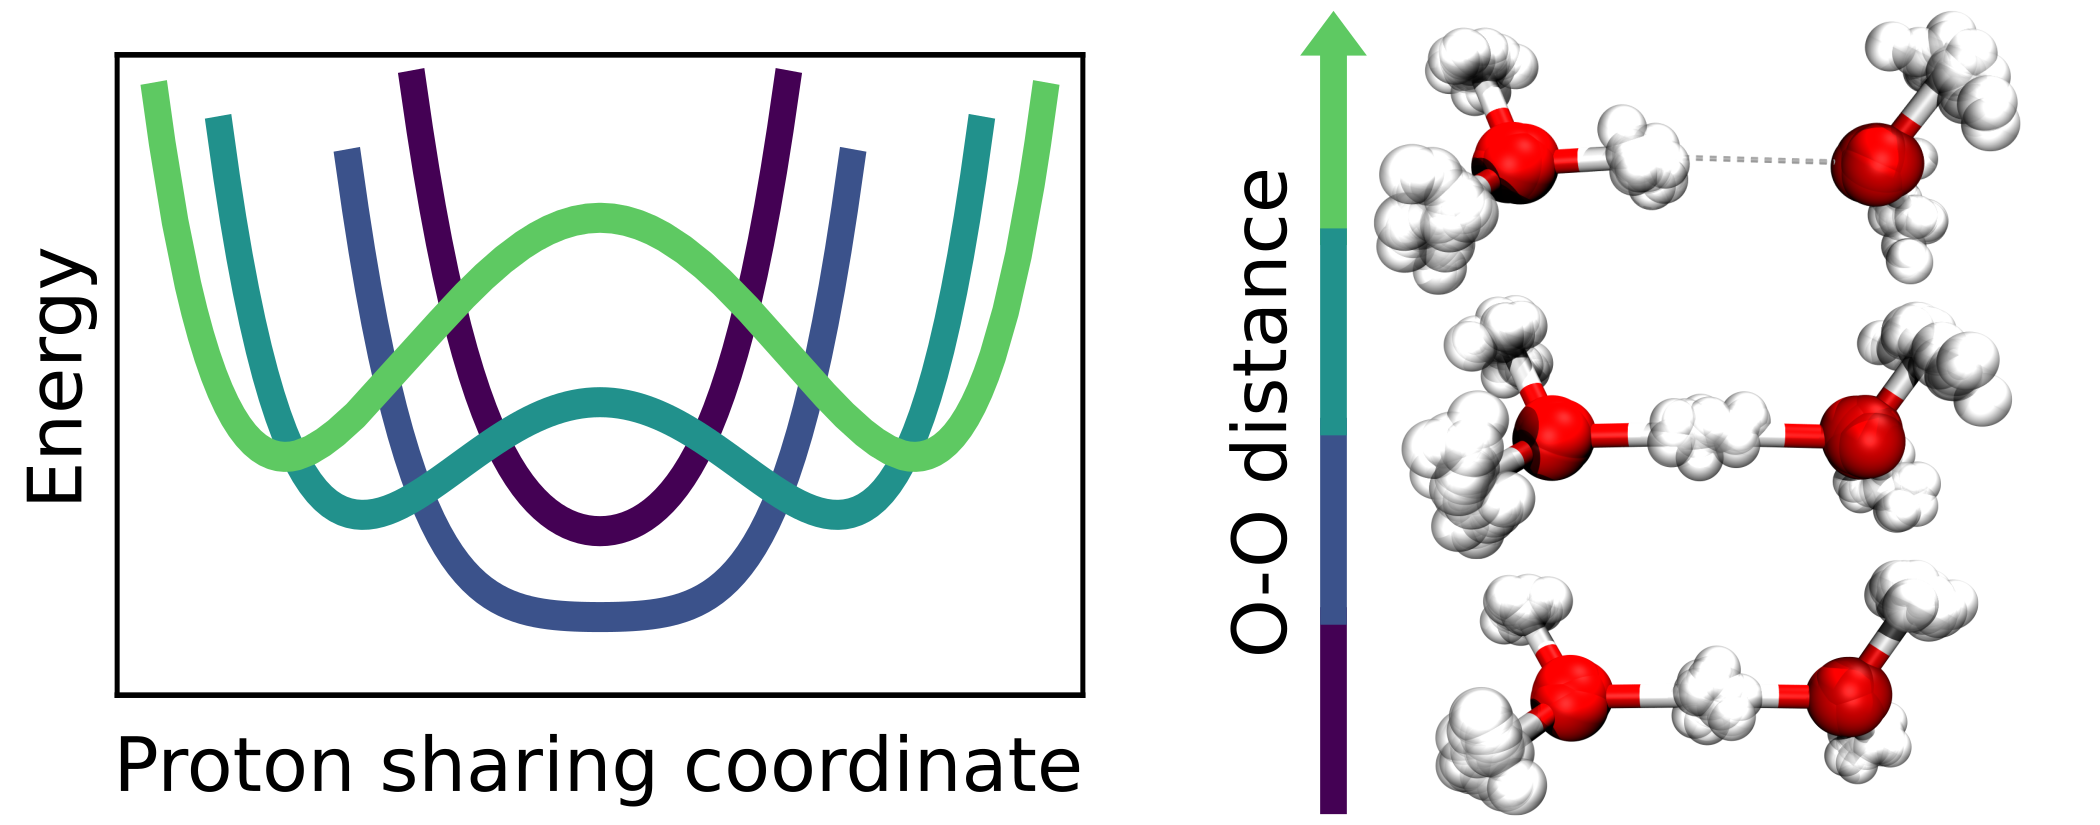
abstract = {For a long time, performing converged path integral simulations at ultra-low, but finite temperatures of a few Kelvin has been a nearly impossible task. However, recent developments in advanced colored noise thermostatting schemes for path integral simulations, namely the Path Integral Generalized Langevin Equation Thermostat (PIGLET) and the Path Integral Quantum Thermal Bath (PIQTB), have been able to greatly reduce the computational cost of these simulations, thus making the ultra-low temperature regime accessible in practice. In this work, we investigate the influence of these two thermostatting schemes on the description of hydrogen-bonded systems at temperatures down to a few Kelvin as encountered, for example, in helium nanodroplet isolation or tagging photodissociation spectroscopy experiments. For this purpose, we analyze the prototypical hydrogen bond in the Zundel cation (H5O2+) as a function of both, oxygen-oxygen distance and temperature in order to elucidate how the anisotropic quantum deloc...},

keywords = {Nuclear Quantum Effects (NQEs), Path Integral Molecular Dynamics (PIMD), Water},

pubstate = {published},

tppubtype = {article}

}
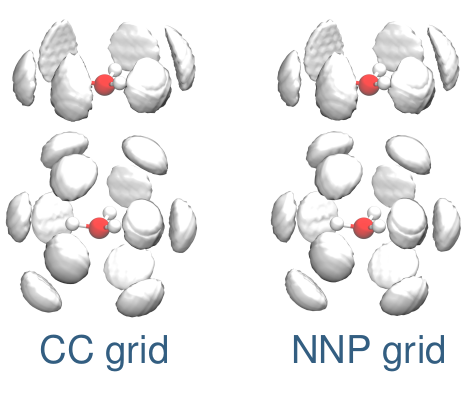
Christoph Schran, Felix Uhl, Jörg Behler, Dominik Marx
High-dimensional neural network potentials for solvation: The case of protonated water clusters in helium Journal Article
In: J. Chem. Phys., vol. 148, no. 10, pp. 102310, 2018, ISSN: 00219606.
Abstract | Links | BibTeX | Tags: Machine Learning Potentials, Nuclear Quantum Effects (NQEs), Superfluidity, Water
@article{Schran2018/10.1063/1.4996819,

title = {High-dimensional neural network potentials for solvation: The case of protonated water clusters in helium},

author = {Christoph Schran and Felix Uhl and Jörg Behler and Dominik Marx},

doi = {10.1063/1.4996819},

issn = {00219606},

year  = {2018},

date = {2018-03-01},

urldate = {2018-03-01},

journal = {J. Chem. Phys.},

volume = {148},

number = {10},

pages = {102310},

abstract = {The design of accurate helium-solute interaction potentials for the simulation of chemically complex molecules solvated in superfluid helium has long been a cumbersome task due to the rather weak but strongly anisotropic nature of the interactions. We show that this challenge can be met by using a combination of an effective pair potential for the He–He interactions and a flexible high-dimensional neural network potential (NNP) for describing the complex interaction between helium and the solute in a pairwise additive manner. This approach yields an excellent agreement with a mean absolute deviation as small as 0.04 kJ mol−1 for the interaction energy between helium and both hydronium and Zundel cations compared with coupled cluster reference calculations with an energetically converged basis set. The construction and improvement of the potential can be performed in a highly automated way, which opens the door for applications to a variety of reactive molecules to study the effect of solvation on the solu...},

keywords = {Machine Learning Potentials, Nuclear Quantum Effects (NQEs), Superfluidity, Water},

pubstate = {published},

tppubtype = {article}

}
2017
Christoph Schran, Ondrej Marsalek, Thomas E. Markland
Unravelling the influence of quantum proton delocalization on electronic charge transfer through the hydrogen bond Journal Article
In: Chem. Phys. Lett., vol. 678, pp. 289–295, 2017, ISSN: 00092614.
Abstract | Links | BibTeX | Tags: Charge Transfer, Hydrogen Bonding, Ions in Water, Nuclear Quantum Effects (NQEs), Water
@article{Schran2017/10.1016/j.cplett.2017.04.034,

title = {Unravelling the influence of quantum proton delocalization on electronic charge transfer through the hydrogen bond},

author = {Christoph Schran and Ondrej Marsalek and Thomas E. Markland},

doi = {10.1016/j.cplett.2017.04.034},

issn = {00092614},

year  = {2017},

date = {2017-06-01},

urldate = {2017-06-01},

journal = {Chem. Phys. Lett.},

volume = {678},

pages = {289–295},

abstract = {Upon hydrogen bond formation, electronic charge density is transferred between the donor and acceptor, impacting processes ranging from hydration to spectroscopy. Here we use ab initio path integral simulations to elucidate the role of nuclear quantum effects in determining the charge transfer in a range of hydrogen bonded species in the gas and liquid phase. We show that the quantization of the nuclei gives rise to large changes in the magnitude of the charge transfer as well as its temperature dependence. We then explain how a single geometric parameter determines the charge transfer through the hydrogen bond. These results thus demonstrate that nuclear quantum effects are vital for the accurate description of charge transfer and offer a physically transparent way to understand how hydrogen bonding gives rise to it.},

keywords = {Charge Transfer, Hydrogen Bonding, Ions in Water, Nuclear Quantum Effects (NQEs), Water},

pubstate = {published},

tppubtype = {article}

}
2016
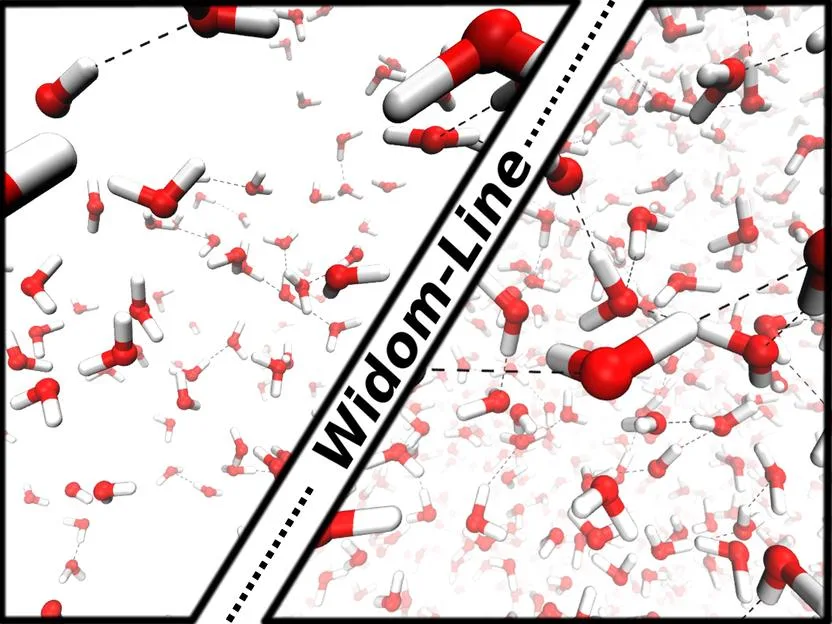
Maciej Śmiechowski, Christoph Schran, Harald Forbert, Dominik Marx
Correlated particle motion and THz spectral response of supercritical water Journal Article
In: Phys. Rev. Lett., vol. 116, no. 2, 2016, ISSN: 10797114.
Abstract | Links | BibTeX | Tags: Hydrogen Bonding, Spectra, Water
@article{Smiechowski2016/10.1103/PhysRevLett.116.027801,

title = {Correlated particle motion and THz spectral response of supercritical water},

author = {Maciej Śmiechowski and Christoph Schran and Harald Forbert and Dominik Marx},

doi = {10.1103/PhysRevLett.116.027801},

issn = {10797114},

year  = {2016},

date = {2016-01-01},

urldate = {2016-01-01},

journal = {Phys. Rev. Lett.},

volume = {116},

number = {2},

abstract = {Molecular dynamics simulations of supercritical water reveal distinctly different distance-dependent modulations of dipolar response and correlations in particle motion compared to ambient conditions. The strongly perturbed H-bond network of water at supercritical conditions allows for considerable translational and rotational freedom of individual molecules. These changes give rise to substantially different infrared spectra and vibrational density of states at THz frequencies for densities above and below the Widom line that separates percolating liquidlike and clustered gaslike supercritical water.},

keywords = {Hydrogen Bonding, Spectra, Water},

pubstate = {published},

tppubtype = {article}

}