
2026

Xavier R. Advincula, Kara D. Fong, Yongkang Wang, Christoph Schran, Mischa Bonn, Angelos Michaelides, Yair Litman
Breaking the air-water paradigm: Ion behavior at hydrophobic solid-water interfaces Journal Article
In: J. Am. Chem. Soc., vol. 148, iss. 12, pp. 12753–12763, 2026.
Abstract | Links | BibTeX | Tags: Hydrogen Bonding, Ions in Water, Machine Learning Potentials, Spectra, Water at Interfaces
@article{Advincula2026/10.1021/jacs.5c20209,

title = {Breaking the air-water paradigm: Ion behavior at hydrophobic solid-water interfaces},

author = {Xavier R. Advincula and Kara D. Fong and Yongkang Wang and Christoph Schran and Mischa Bonn and Angelos Michaelides and Yair Litman},

url = {https://pubs.acs.org/doi/full/10.1021/jacs.5c20209},

doi = {10.1021/jacs.5c20209},

year  = {2026},

date = {2026-03-19},

journal = {J. Am. Chem. Soc.},

volume = {148},

issue = {12},

pages = {12753–12763},

abstract = {Hydrophobic solid-water interfaces underpin processes in nanofluidics, electrochemistry, and energy technologies. Microscopic insights into these systems are often inferred from our understanding of the air-water interface, which is assumed to exhibit similar behavior. Here, we challenge this paradigm by combining heterodyne-detected vibrational sum-frequency generation spectroscopy with machine-learning molecular dynamics simulations at first-principles accuracy to investigate the graphene-NaCl(aq) interface as a prototypical hydrophobic solid-water system. Spectroscopic results suggest that ions have a minimal effect on the structure of the interfacial water, while simulations reveal that Na+ and Cl- accumulate densely at the surface. Together, these findings reveal a new adsorption mechanism that departs from the established air-water interface paradigm, where interfacial ion adsorption is typically associated with, and often detected through, a pronounced alteration of the interfacial water alignment and orientation. This difference arises because ions cannot penetrate the solid boundary and reside at a similar depth as the interfacial water molecules. As a consequence, large ion populations can be accommodated within the extended two-dimensional hydrogen-bond network at the interface, causing only minor local distortions but significant changes to its longer-range connectivity. These results reveal a distinct mechanism of electrolyte organization at aqueous-carbon interfaces, relevant to energy applications, where performance is highly sensitive to the local organization of interfacial water.},

keywords = {Hydrogen Bonding, Ions in Water, Machine Learning Potentials, Spectra, Water at Interfaces},

pubstate = {published},

tppubtype = {article}

}

Yixuan Feng, Xavier R. Advincula, Hongwei Fang, Christoph Schran
Clay edges are dynamic proton-conducting networks modulated by structure and pH Journal Article
In: J. Phys. Chem. Lett., vol. 17, iss. 9, pp. 2679–2688, 2026.
Abstract | Links | BibTeX | Tags: Charge Transfer, Chemical Reactivity, Ions in Water, Water, Water at Interfaces
@article{Feng2026/10.1021/acs.jpclett.5c03748,

title = {Clay edges are dynamic proton-conducting networks modulated by structure and pH},

author = {Yixuan Feng and Xavier R. Advincula and Hongwei Fang and Christoph Schran},

url = {https://pubs.acs.org/doi/full/10.1021/acs.jpclett.5c03748},

doi = {10.1021/acs.jpclett.5c03748},

year  = {2026},

date = {2026-02-22},

urldate = {2026-02-22},

journal = {J. Phys. Chem. Lett.},

volume = {17},

issue = {9},

pages = {2679–2688},

abstract = {Montmorillonite, a ubiquitous clay mineral, plays a vital role in geochemical and environmental processes due to its chemically complex edge surfaces. However, the molecular-scale acid-base reactivity of these interfaces remains poorly understood due to the limitations of both experimental resolution and conventional simulations. Here, we employ machine learning potentials with first-principles accuracy to perform nanosecond-scale molecular dynamics simulations of montmorillonite nanoparticles across a range of pH. Our results reveal clear amphoteric behavior, with edge sites undergoing protonation in acidic environments and deprotonation in basic conditions, accompanied by pH-dependent surface charge regulation. Even at neutral pH, spontaneous and directional proton transfer events are common, proceeding via both direct and solvent-mediated pathways. These findings demonstrate that montmorillonite edges are not static arrays of hydroxyl groups but dynamic, proton-conducting networks whose reactivity and charge state are modulated by local structure and solution conditions. This work offers a molecular-level framework for understanding proton transport and buffering in clay-water systems, with broad implications for catalysis, ion exchange, and environmental remediation.},

keywords = {Charge Transfer, Chemical Reactivity, Ions in Water, Water, Water at Interfaces},

pubstate = {published},

tppubtype = {article}

}
2024

Niamh O'Neill, Christoph Schran, Stephen J. Cox, Angelos Michaelides
Crumbling crystals: on the dissolution mechanism of NaCl in water Journal Article
In: Phys. Chem. Chem. Phys., vol. 26, iss. 42, pp. 26933-26942, 2024.
Abstract | Links | BibTeX | Tags: Ions in Water, Machine Learning Potentials, Water
@article{ONeill2024/10.1039/d4cp03115f,

title = {Crumbling crystals: on the dissolution mechanism of NaCl in water},

author = {Niamh O'Neill and Christoph Schran and Stephen J. Cox and Angelos Michaelides},

url = {https://pubs.rsc.org/en/content/articlelanding/2024/cp/d4cp03115f},

doi = {10.1039/d4cp03115f},

year  = {2024},

date = {2024-10-30},

urldate = {2024-10-30},

journal = {Phys. Chem. Chem. Phys.},

volume = {26},

issue = {42},

pages = {26933-26942},

abstract = {Dissolution of ionic salts in water is ubiquitous, particularly for NaCl. However, an atomistic scale understanding of the process remains elusive. Simulations lend themselves conveniently to studying dissolution since they provide the spatio-temporal resolution that can be difficult to obtain experimentally. Nevertheless, the complexity of various inter- and intra-molecular interactions require careful treatment and long time scale simulations, both of which are typically hindered by computational expense. Here, we use advances in machine learning potential methodology to resolve at an ab initio level of theory the dissolution mechanism of NaCl in water. The picture that emerges is that of a steady ion-wise unwrapping of the crystal preceding its rapid disintegration, reminiscent of crumbling. The onset of crumbling can be explained by a strong increase in the ratio of the surface area to volume of the crystal. Overall, dissolution comprises a series of highly dynamical microscopic sub-processes, resulting in an inherently stochastic mechanism. These atomistic level insights contribute to the general understanding of dissolution mechanisms in other crystals, and the methodology is primed for more complex systems of recent interest such as water/salt interfaces under flow and salt crystals under confinement.},

keywords = {Ions in Water, Machine Learning Potentials, Water},

pubstate = {published},

tppubtype = {article}

}
Niamh O’Neill, Benjamin X. Shi, Kara D. Fong, Angelos Michaelides, Christoph Schran
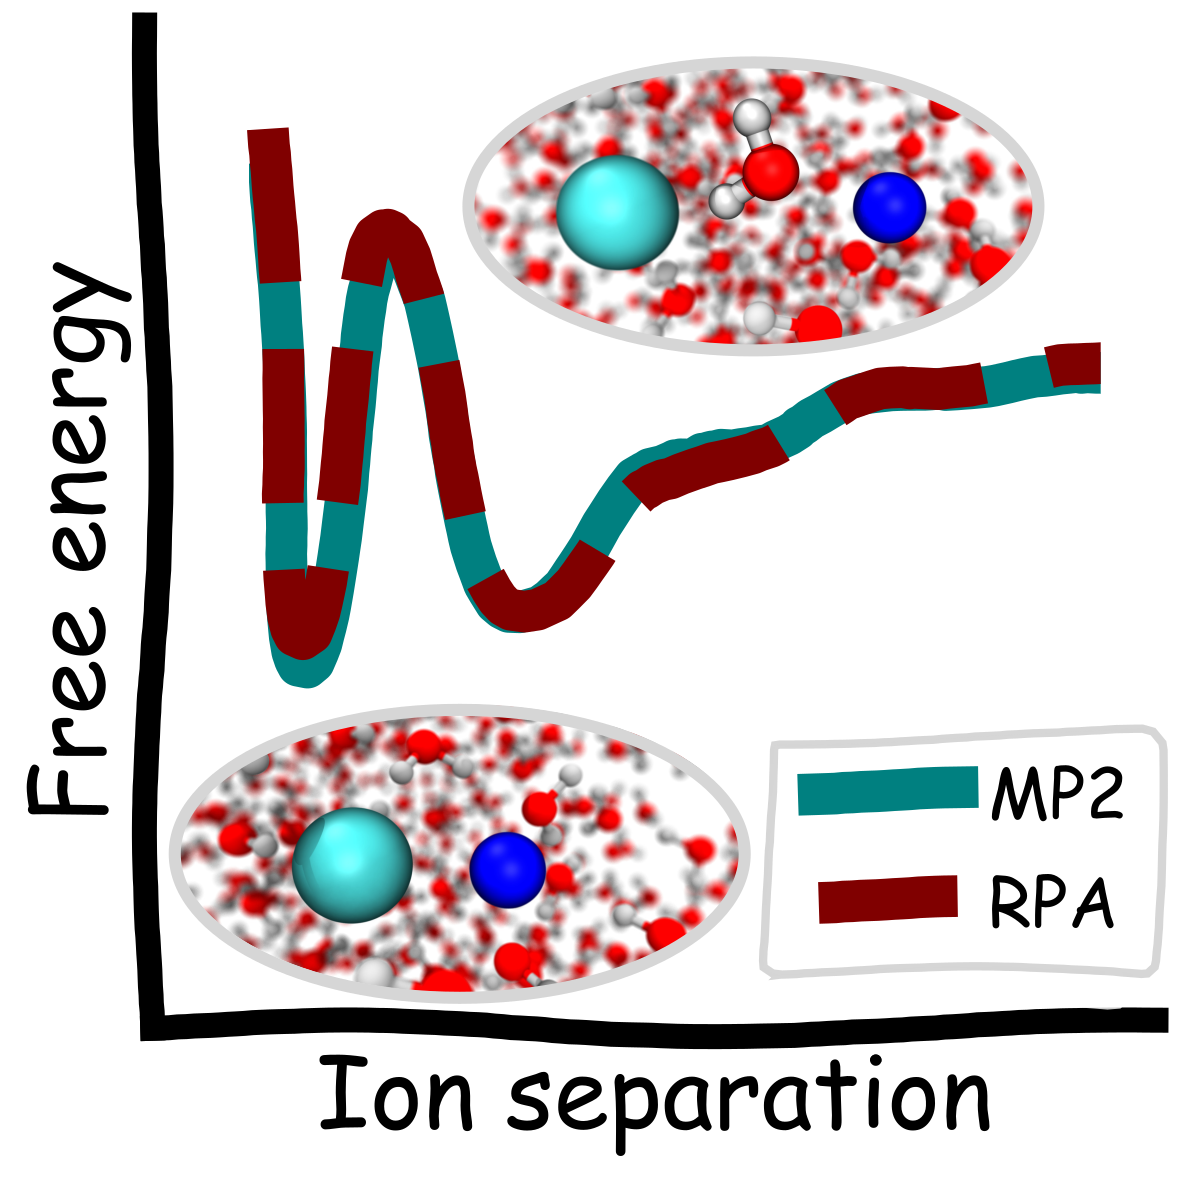
To pair or not to pair? Machine-learned explicitly-correlated electronic structure for NaCl in water Journal Article
In: The Journal of Physical Chemistry Letters, vol. 15, no. 23, pp. 6081–6091, 2024, (PMID: 38820256).
Abstract | Links | BibTeX | Tags: Ions in Water, Machine Learning Potentials
@article{ONeill/10.1021/acs.jpclett.4c01030,

title = {To pair or not to pair? Machine-learned explicitly-correlated electronic structure for NaCl in water},

author = {Niamh O’Neill and Benjamin X. Shi and Kara D. Fong and Angelos Michaelides and Christoph Schran},

url = {https://doi.org/10.1021/acs.jpclett.4c01030},

doi = {10.1021/acs.jpclett.4c01030},

year  = {2024},

date = {2024-05-31},

urldate = {2024-05-31},

journal = {The Journal of Physical Chemistry Letters},

volume = {15},

number = {23},

pages = {6081–6091},

abstract = {The extent of ion pairing in solution is an important phenomenon to rationalize transport and thermodynamic properties of electrolytes. A fundamental measure of this pairing is the potential of mean force (PMF) between solvated ions. The relative stabilities of the paired and solvent shared states in the PMF and the barrier between them are highly sensitive to the underlying potential energy surface. However, direct application of accurate electronic structure methods is challenging, since long simulations are required. We develop wave function based machine learning potentials with the random phase approximation (RPA) and second order Møller–Plesset (MP2) perturbation theory for the prototypical system of Na and Cl ions in water. We show both methods in agreement, predicting the paired and solvent shared states to have similar energies (within 0.2 kcal/mol). We also provide the same benchmarks for different DFT functionals as well as insight into the PMF based on simple analyses of the interactions in the system.},

note = {PMID: 38820256},

keywords = {Ions in Water, Machine Learning Potentials},

pubstate = {published},

tppubtype = {article}

}
Kara D. Fong, Barbara Sumić, Niamh O’Neill, Christoph Schran, Clare P. Grey, Angelos Michaelides
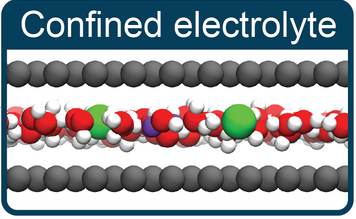
The interplay of solvation and polarization effects on ion pairing in nanoconfined electrolytes Journal Article
In: Nano Letters, vol. 24, no. 16, pp. 5024–5030, 2024, (PMID: 38592099).
Abstract | Links | BibTeX | Tags: Confinement, Hydrogen Bonding, Ions in Water, Machine Learning Potentials
@article{Fong/10.1021/acs.nanolett.4c00890,

title = {The interplay of solvation and polarization effects on ion pairing in nanoconfined electrolytes},

author = {Kara D. Fong and Barbara Sumić and Niamh O’Neill and Christoph Schran and Clare P. Grey and Angelos Michaelides},

url = {https://doi.org/10.1021/acs.nanolett.4c00890},

doi = {10.1021/acs.nanolett.4c00890},

year  = {2024},

date = {2024-04-09},

urldate = {2024-04-09},

journal = {Nano Letters},

volume = {24},

number = {16},

pages = {5024–5030},

abstract = {The nature of ion–ion interactions in electrolytes confined to nanoscale pores has important implications for energy storage and separation technologies. However, the physical effects dictating the structure of nanoconfined electrolytes remain debated. Here we employ machine-learning-based molecular dynamics simulations to investigate ion–ion interactions with density functional theory level accuracy in a prototypical confined electrolyte, aqueous NaCl within graphene slit pores. We find that the free energy of ion pairing in highly confined electrolytes deviates substantially from that in bulk solutions, observing a decrease in contact ion pairing but an increase in solvent-separated ion pairing. These changes arise from an interplay of ion solvation effects and graphene’s electronic structure. Notably, the behavior observed from our first-principles-level simulations is not reproduced even qualitatively with the classical force fields conventionally used to model these systems. The insight provided in this work opens new avenues for predicting and controlling the structure of nanoconfined electrolytes.},

note = {PMID: 38592099},

keywords = {Confinement, Hydrogen Bonding, Ions in Water, Machine Learning Potentials},

pubstate = {published},

tppubtype = {article}

}
2021
Christoph Schran, Fabian L. Thiemann, Patrick Rowe, Erich A. Müller, Ondrej Marsalek, Angelos Michaelides
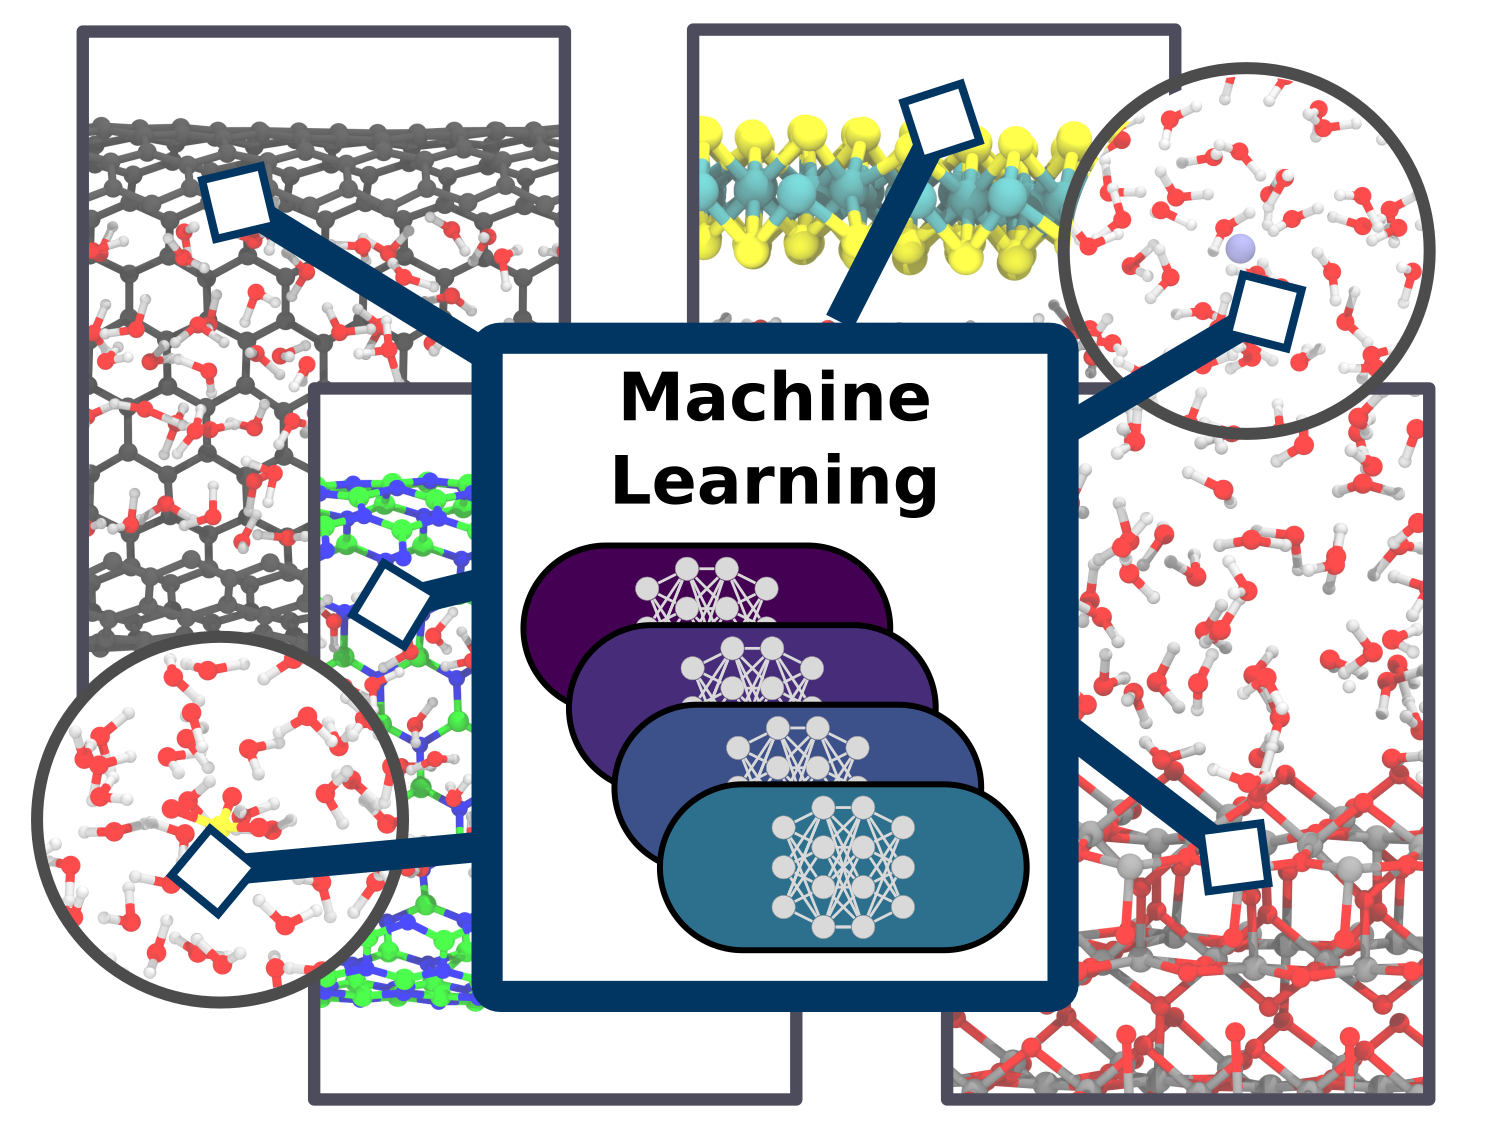
Machine learning potentials for complex aqueous systems made simple Journal Article
In: Proc. Natl. Acad. Sci., vol. 118, no. 38, pp. e2110077118, 2021, ISSN: 0027-8424.
Abstract | Links | BibTeX | Tags: Confinement, Ions in Water, Machine Learning Potentials, Water, Water at Interfaces
@article{Schran2021/10.1073/PNAS.2110077118,

title = {Machine learning potentials for complex aqueous systems made simple},

author = {Christoph Schran and Fabian L. Thiemann and Patrick Rowe and Erich A. Müller and Ondrej Marsalek and Angelos Michaelides},

doi = {10.1073/PNAS.2110077118},

issn = {0027-8424},

year  = {2021},

date = {2021-09-01},

urldate = {2021-09-01},

journal = {Proc. Natl. Acad. Sci.},

volume = {118},

number = {38},

pages = {e2110077118},

publisher = {National Academy of Sciences},

abstract = {Simulation techniques based on accurate and efficient representations of potential energy surfaces are urgently needed for the understanding of complex aqueous systems such as solid-liquid interfaces. Here, we present a machine learning framework that enables the efficient development and validation of models for complex aqueous systems. Instead of trying to deliver a globally-optimal machine learning potential, we propose to develop models applicable to specific thermodynamic state points in a simple and user-friendly process. After an initial ab initio simulation, a machine learning potential is constructed with minimum human effort through a data-driven active learning protocol. Such models can afterwards be applied in exhaustive simulations to provide reliable answers for the scientific question at hand. We showcase this methodology on a diverse set of aqueous systems with increasing degrees of complexity. The systems chosen here comprise water with different ions in solution, water on a titanium dioxide surface, as well as water confined in nanotubes and between molybdenum disulfide sheets. Highlighting the accuracy of our approach compared to the ab initio reference, the resulting models are evaluated in detail with an automated validation protocol that includes structural and dynamical properties and the precision of the force prediction of the models. Finally, we demonstrate the capabilities of our approach for the description of water on the rutile titanium dioxide (110) surface to analyze structure and mobility of water on this surface. Such machine learning models provide a straightforward and uncomplicated but accurate extension of simulation time and length scales for complex systems.},

keywords = {Confinement, Ions in Water, Machine Learning Potentials, Water, Water at Interfaces},

pubstate = {published},

tppubtype = {article}

}
2017
Christoph Schran, Ondrej Marsalek, Thomas E. Markland
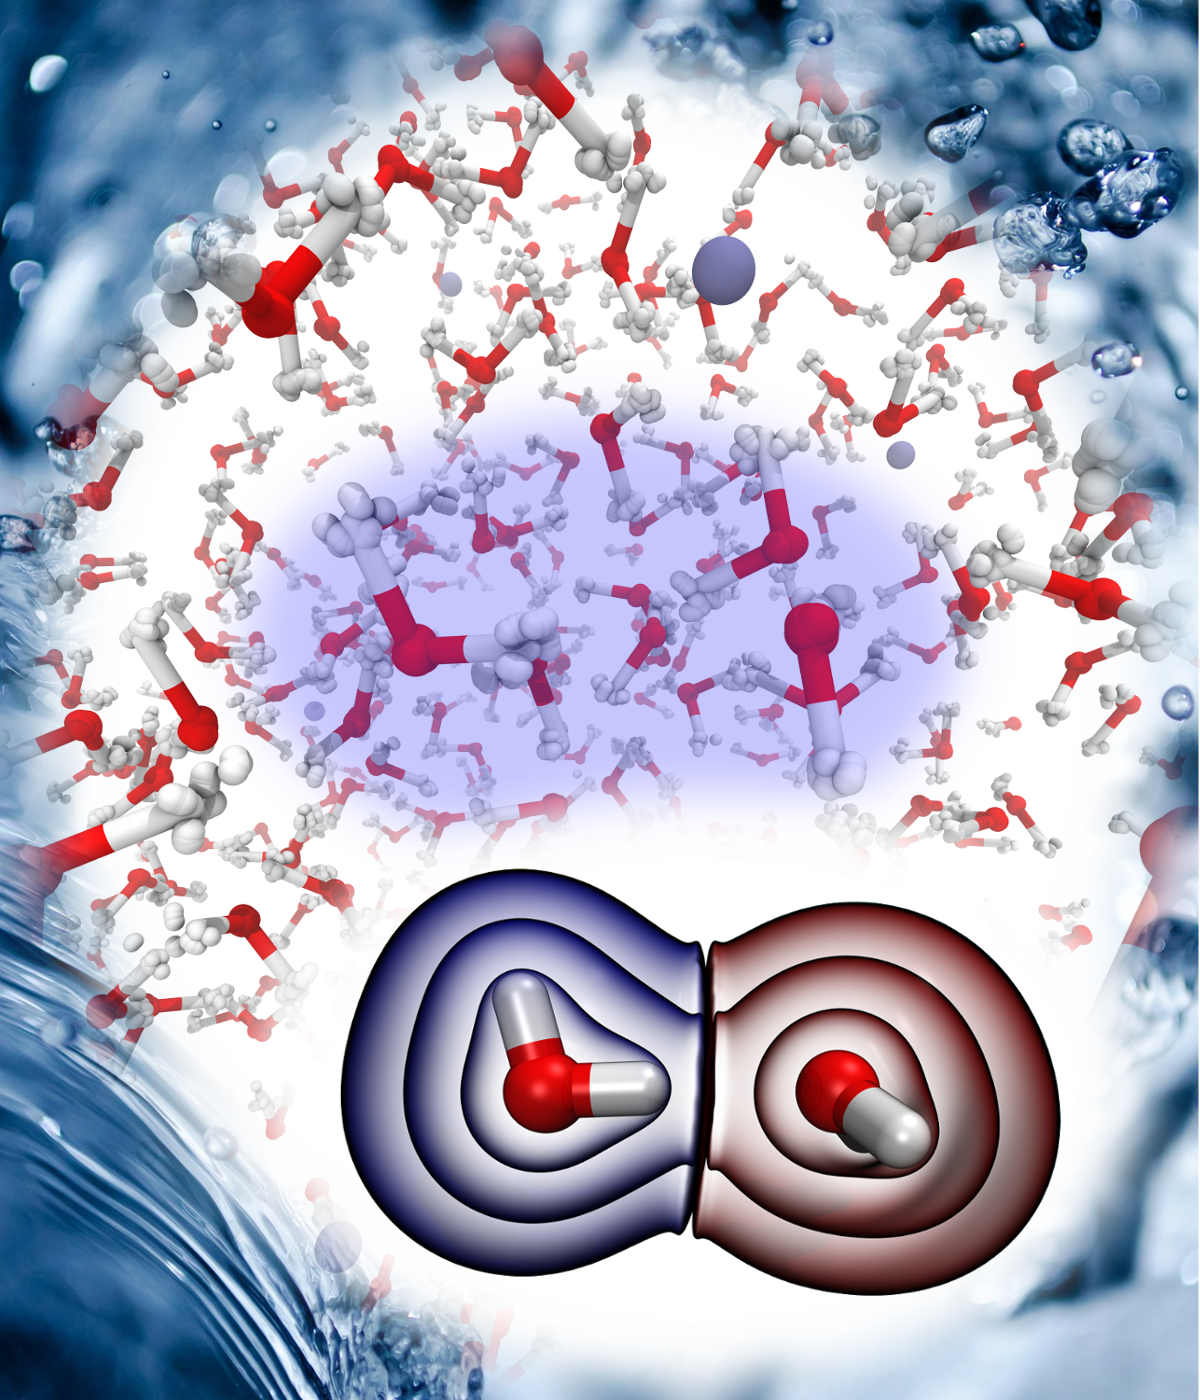
Unravelling the influence of quantum proton delocalization on electronic charge transfer through the hydrogen bond Journal Article
In: Chem. Phys. Lett., vol. 678, pp. 289–295, 2017, ISSN: 00092614.
Abstract | Links | BibTeX | Tags: Charge Transfer, Hydrogen Bonding, Ions in Water, Nuclear Quantum Effects (NQEs), Water
@article{Schran2017/10.1016/j.cplett.2017.04.034,

title = {Unravelling the influence of quantum proton delocalization on electronic charge transfer through the hydrogen bond},

author = {Christoph Schran and Ondrej Marsalek and Thomas E. Markland},

doi = {10.1016/j.cplett.2017.04.034},

issn = {00092614},

year  = {2017},

date = {2017-06-01},

urldate = {2017-06-01},

journal = {Chem. Phys. Lett.},

volume = {678},

pages = {289–295},

abstract = {Upon hydrogen bond formation, electronic charge density is transferred between the donor and acceptor, impacting processes ranging from hydration to spectroscopy. Here we use ab initio path integral simulations to elucidate the role of nuclear quantum effects in determining the charge transfer in a range of hydrogen bonded species in the gas and liquid phase. We show that the quantization of the nuclei gives rise to large changes in the magnitude of the charge transfer as well as its temperature dependence. We then explain how a single geometric parameter determines the charge transfer through the hydrogen bond. These results thus demonstrate that nuclear quantum effects are vital for the accurate description of charge transfer and offer a physically transparent way to understand how hydrogen bonding gives rise to it.},

keywords = {Charge Transfer, Hydrogen Bonding, Ions in Water, Nuclear Quantum Effects (NQEs), Water},

pubstate = {published},

tppubtype = {article}

}