
2026

Darren Wayne Lim, Xavier R. Advincula, William C. Witt, Angelos Michaelides, Fabian L. Thiemann, Christoph Schran
Strain-dependent wetting of graphene Journal Article
In: Chem. Sci., 2026.
Abstract | Links | BibTeX | Tags: Machine Learning Potentials, Two-dimensional Materials, Water, Water at Interfaces
@article{Lim2026/10.1039/d6sc05377g,

title = {Strain-dependent wetting of graphene},

author = {Darren Wayne Lim and Xavier R. Advincula and William C. Witt and Angelos Michaelides and Fabian L. Thiemann and Christoph Schran},

url = {https://doi.org/10.1039/d6sc05377g},

doi = {10.1039/d6sc05377g},

year  = {2026},

date = {2026-10-20},

urldate = {2026-10-20},

journal = {Chem. Sci.},

abstract = {Understanding how water wets graphene is critical for predicting and controlling its behaviour in nanofluidic, sensing, and energy applications. A key measure of wetting is the contact angle made by a liquid droplet against the surface, yet experimental measurements for graphene span a wide range, with no consensus for free-standing graphene. Here, we use a machine learning potential with ab initio accuracy to provide an atomistic first-principles prediction for this unsolved problem, finding a weakly hydrophilic contact angle of 72.1 ± 1.5°. More importantly, we unveil that graphene's wetting properties are highly sensitive to mechanical strain: tensile strain makes graphene significantly less hydrophilic, while compressive strain induces coherent ripples around the droplet, resulting in pronounced anisotropic wetting and contact angle hysteresis. We show that there is a strong coupling between the three-phase contact line and the intrinsic thermal ripples of free-standing graphene, which contributes to this strain sensitivity. Our results demonstrate that the wettability of 2D membranes is governed not only by their chemistry but also by their dynamic morphology, introducing a new class of wetting behaviour unique to atomically thin materials that offers an additional explanation for variability in experimental measurements. These findings reveal that mechanical strain may be a practical route to controlling wetting in 2D nanomaterials-based technologies, with promising consequences for nanofluidic and nano-filtration applications.},

keywords = {Machine Learning Potentials, Two-dimensional Materials, Water, Water at Interfaces},

pubstate = {published},

tppubtype = {article}

}

Xavier R. Advincula, Yair Litman, Kara D. Fong, William C. Witt, Christoph Schran, Angelos Michaelides
How reactive is water at the nanoscale and how to control it? Journal Article
In: Sci. Adv., vol. 12, iss. 26, pp. eaeb5772, 2026.
Abstract | Links | BibTeX | Tags: Chemical Reactivity, Confinement, Machine Learning Potentials, Two-dimensional Materials, Water, Water at Interfaces
@article{Advincula2026/10.1126/sciadv.aeb5772,

title = {How reactive is water at the nanoscale and how to control it?},

author = {Xavier R. Advincula and Yair Litman and Kara D. Fong and William C. Witt and Christoph Schran and Angelos Michaelides},

url = {https://www.science.org/doi/10.1126/sciadv.aeb5772},

doi = {10.1126/sciadv.aeb5772},

year  = {2026},

date = {2026-06-24},

journal = {Sci. Adv.},

volume = {12},

issue = {26},

pages = {eaeb5772},

abstract = {Nanoconfined water plays a key role in nanofluidics, electrochemistry, and catalysis, yet its reactivity remains a matter of debate. Prior studies have reported both enhanced and suppressed water self-dissociation relative to the bulk, but without a consistent explanation. Here, using enhanced sampling molecular dynamics with machine-learned potentials trained at first-principles accuracy, we investigate dissociation behavior in water confined within two-dimensional slit pores and nanodroplets, using graphene and hexagonal boron nitride as model materials. We find that reactivity is extremely sensitive to water density, geometry, and surface chemistry, among other factors. Despite this complexity, we show that chemical potential, together with interfacial interactions, governs dissociation trends and explains the variability observed in prior studies. Within this framework, when confined water is compared to the bulk at equivalent chemical potential, corresponding to thermodynamic equilibrium with a bulk reservoir, its reactivity remains essentially unchanged; rather, differences arise when the systems are compared at different chemical potentials or under distinct interfacial conditions. This thermodynamic perspective reconciles previous contradictions and reveals how nanoscale environments can drastically shift water reactivity. Our findings provide molecular-level insight and offer a design lever for modulating water chemistry at the nanoscale.},

keywords = {Chemical Reactivity, Confinement, Machine Learning Potentials, Two-dimensional Materials, Water, Water at Interfaces},

pubstate = {published},

tppubtype = {article}

}

Xavier R. Advincula, Kara D. Fong, Yongkang Wang, Christoph Schran, Mischa Bonn, Angelos Michaelides, Yair Litman
Breaking the air-water paradigm: Ion behavior at hydrophobic solid-water interfaces Journal Article
In: J. Am. Chem. Soc., vol. 148, iss. 12, pp. 12753–12763, 2026.
Abstract | Links | BibTeX | Tags: Hydrogen Bonding, Ions in Water, Machine Learning Potentials, Spectra, Water at Interfaces
@article{Advincula2026/10.1021/jacs.5c20209,

title = {Breaking the air-water paradigm: Ion behavior at hydrophobic solid-water interfaces},

author = {Xavier R. Advincula and Kara D. Fong and Yongkang Wang and Christoph Schran and Mischa Bonn and Angelos Michaelides and Yair Litman},

url = {https://pubs.acs.org/doi/full/10.1021/jacs.5c20209},

doi = {10.1021/jacs.5c20209},

year  = {2026},

date = {2026-03-19},

journal = {J. Am. Chem. Soc.},

volume = {148},

issue = {12},

pages = {12753–12763},

abstract = {Hydrophobic solid-water interfaces underpin processes in nanofluidics, electrochemistry, and energy technologies. Microscopic insights into these systems are often inferred from our understanding of the air-water interface, which is assumed to exhibit similar behavior. Here, we challenge this paradigm by combining heterodyne-detected vibrational sum-frequency generation spectroscopy with machine-learning molecular dynamics simulations at first-principles accuracy to investigate the graphene-NaCl(aq) interface as a prototypical hydrophobic solid-water system. Spectroscopic results suggest that ions have a minimal effect on the structure of the interfacial water, while simulations reveal that Na+ and Cl- accumulate densely at the surface. Together, these findings reveal a new adsorption mechanism that departs from the established air-water interface paradigm, where interfacial ion adsorption is typically associated with, and often detected through, a pronounced alteration of the interfacial water alignment and orientation. This difference arises because ions cannot penetrate the solid boundary and reside at a similar depth as the interfacial water molecules. As a consequence, large ion populations can be accommodated within the extended two-dimensional hydrogen-bond network at the interface, causing only minor local distortions but significant changes to its longer-range connectivity. These results reveal a distinct mechanism of electrolyte organization at aqueous-carbon interfaces, relevant to energy applications, where performance is highly sensitive to the local organization of interfacial water.},

keywords = {Hydrogen Bonding, Ions in Water, Machine Learning Potentials, Spectra, Water at Interfaces},

pubstate = {published},

tppubtype = {article}

}

Yixuan Feng, Xavier R. Advincula, Hongwei Fang, Christoph Schran
Clay edges are dynamic proton-conducting networks modulated by structure and pH Journal Article
In: J. Phys. Chem. Lett., vol. 17, iss. 9, pp. 2679–2688, 2026.
Abstract | Links | BibTeX | Tags: Charge Transfer, Chemical Reactivity, Ions in Water, Water, Water at Interfaces
@article{Feng2026/10.1021/acs.jpclett.5c03748,

title = {Clay edges are dynamic proton-conducting networks modulated by structure and pH},

author = {Yixuan Feng and Xavier R. Advincula and Hongwei Fang and Christoph Schran},

url = {https://pubs.acs.org/doi/full/10.1021/acs.jpclett.5c03748},

doi = {10.1021/acs.jpclett.5c03748},

year  = {2026},

date = {2026-02-22},

urldate = {2026-02-22},

journal = {J. Phys. Chem. Lett.},

volume = {17},

issue = {9},

pages = {2679–2688},

abstract = {Montmorillonite, a ubiquitous clay mineral, plays a vital role in geochemical and environmental processes due to its chemically complex edge surfaces. However, the molecular-scale acid-base reactivity of these interfaces remains poorly understood due to the limitations of both experimental resolution and conventional simulations. Here, we employ machine learning potentials with first-principles accuracy to perform nanosecond-scale molecular dynamics simulations of montmorillonite nanoparticles across a range of pH. Our results reveal clear amphoteric behavior, with edge sites undergoing protonation in acidic environments and deprotonation in basic conditions, accompanied by pH-dependent surface charge regulation. Even at neutral pH, spontaneous and directional proton transfer events are common, proceeding via both direct and solvent-mediated pathways. These findings demonstrate that montmorillonite edges are not static arrays of hydroxyl groups but dynamic, proton-conducting networks whose reactivity and charge state are modulated by local structure and solution conditions. This work offers a molecular-level framework for understanding proton transport and buffering in clay-water systems, with broad implications for catalysis, ion exchange, and environmental remediation.},

keywords = {Charge Transfer, Chemical Reactivity, Ions in Water, Water, Water at Interfaces},

pubstate = {published},

tppubtype = {article}

}

Xavier R. Advincula, Christoph Schran, Angelos Michaelides
When is nanoconfined water different from interfacial water? Journal Article
In: Faraday Discuss., 2026.
Abstract | Links | BibTeX | Tags: Confinement, Hydrogen Bonding, Two-dimensional Materials, Water, Water at Interfaces
@article{Advincula2026/10.1039/d5fd00165j,

title = {When is nanoconfined water different from interfacial water?},

author = {Xavier R. Advincula and Christoph Schran and Angelos Michaelides},

url = {https://pubs.rsc.org/en/content/articlelanding/2026/fd/d5fd00165j},

doi = {10.1039/D5FD00165J},

year  = {2026},

date = {2026-01-14},

urldate = {2026-01-14},

journal = {Faraday Discuss.},

abstract = {Water behaves very differently at surfaces and under extreme confinement, but the boundary between these two regimes has remained unclear. Despite evidence that interfacial effects persist under sub-nanometre confinement, the molecular-scale behaviour and its evolution with slit width remain unclear. Here, we use machine-learning molecular dynamics with first-principles accuracy to probe water at graphene surfaces across slit widths ranging from the open-interface limit to angstrom-scale confinement. We find that water undergoes a sharp structural transition: when three or more water layers fit between the walls, the structure of the graphene-water interface is effectively indistinguishable from that in an open system, with density layering, hydrogen bonding, and orientational ordering retaining interfacial character. Below this threshold, however, angstrom-scale confinement strongly reorganises the liquid, producing enhanced ordering, a restructured hydrogen-bond network, and modified orientational motifs. These results establish a molecular-level picture that clearly separates interfacial behaviour from genuine nanoconfinement and provide guidance for predicting and controlling the structure of water in nanoscale solid-liquid environments.},

keywords = {Confinement, Hydrogen Bonding, Two-dimensional Materials, Water, Water at Interfaces},

pubstate = {published},

tppubtype = {article}

}
2025

Zeke Coady, Samuel G.H. Brookes, Zhaohan Shen, Benjamin J. Rhodes, Grace Mapstone, Zhen Xu, Wei Yu, Hirotomo Nishihara, Christoph Schran, Angelos Michaelides, Alexander C. Forse
Unexpected oversolubility of CO₂ measured at electrode–electrolyte interfaces Journal Article
In: J. Am. Chem. Soc., vol. 147, iss. 40, pp. 36310–36319, 2025.
Abstract | Links | BibTeX | Tags: Chemical Reactivity, Machine Learning Potentials, Water at Interfaces
@article{Coady2025/10.1021/jacs.5c09712,

title = {Unexpected oversolubility of CO₂ measured at electrode–electrolyte interfaces},

author = {Zeke Coady and Samuel G.H. Brookes and Zhaohan Shen and Benjamin J. Rhodes and Grace Mapstone and Zhen Xu and Wei Yu and Hirotomo Nishihara and Christoph Schran and Angelos Michaelides and Alexander C. Forse},

url = {https://pubs.acs.org/doi/full/10.1021/jacs.5c09712},

doi = {10.1021/jacs.5c09712},

year  = {2025},

date = {2025-09-23},

urldate = {2025-09-23},

journal = {J. Am. Chem. Soc.},

volume = {147},

issue = {40},

pages = {36310–36319},

abstract = {Enhancements in gas solubility in pore-confined liquids─termed oversolubility─can drastically influence gas separation and catalytic efficiency in confined environments; however, they remain poorly understood in electrochemical $CO_2$ capture and reduction systems. While previous investigations of oversolubility have emphasized the importance of mesoporosity and incomplete pore saturation by the solvent, in this work, we report an unprecedented 30-fold oversolubility effect for $CO_2$ in solely microporous activated carbons saturated with 1 M $Na_2SO_{4(aq)}$. The oversolubility effect occurs regardless of the activated carbon’s functional groups and level of disorder and is enhanced for smaller pore sizes. Oversolubility is quantified using solid-state $^{13}C$ nuclear magnetic resonance spectroscopy (NMR), enabling differentiation between in-pore and ex-pore $CO_2$ and $HCO_3^–$. Atomistic modeling of the system, based on a machine-learning model delivering first-principles accuracy, suggests that the effect is driven by an adsorption-like mechanism underpinned by favorable interactions between $CO_2$ and the pore walls. Our findings demonstrate the unexpected importance of oversolubility for gas uptake in microporous, solvent-saturated carbon electrodes, an effect with direct relevance for improving electrochemical $CO_2$ capture and conversion technologies.},

keywords = {Chemical Reactivity, Machine Learning Potentials, Water at Interfaces},

pubstate = {published},

tppubtype = {article}

}

Samuel G.H. Brookes, Venkat Kapil, Angelos Michaelides, Christoph Schran
CO₂ hydration at the air–water interface: A surface-mediated “in-and-out” mechanism Journal Article
In: Proc. Natl. Acad. Sci., vol. 122, no. 34, pp. e2502684122, 2025.
Abstract | Links | BibTeX | Tags: Chemical Reactivity, Machine Learning Potentials, Water at Interfaces
@article{Brookes2025/10.1073/pnas.2502684122,

title = {CO₂ hydration at the air–water interface: A surface-mediated “in-and-out” mechanism},

author = {Samuel G.H. Brookes and Venkat Kapil and Angelos Michaelides and Christoph Schran},

url = {https://www.pnas.org/doi/full/10.1073/pnas.2502684122},

doi = {10.1073/pnas.2502684122},

year  = {2025},

date = {2025-08-20},

urldate = {2025-08-20},

journal = {Proc. Natl. Acad. Sci.},

volume = {122},

number = {34},

pages = {e2502684122},

abstract = {An understanding of the $CO_2 + H_2O$ hydration reaction is crucial for modeling the effects of ocean acidification, for enabling novel carbon storage solutions, and as a model process in the geosciences. While the mechanism of this reaction has been investigated extensively in the condensed phase, its mechanism at the air–water interface remains elusive, leaving uncertain the contribution that surface-adsorbed $CO_2$ makes to the overall acidification reaction. In this study, we employ machine-learned potentials trained to various levels of theory to provide a molecular-level understanding of $CO_2$ hydration at the air–water interface. We show that reaction at the interface follows a surface-mediated “in-and-out” mechanism: $CO_2$ diffuses into the aqueous surface layer, reacts to form carbonic acid, and is subsequently expelled from solution. We show that this surface layer provides a bulk-like solvation environment, engendering similar modes of reactivity and near-identical free energy profiles for the bulk and interfacial processes. Our study unveils an unconventional reaction mechanism that underscores the dynamic nature of the molecular reaction site at the air–water interface. The similarity between bulk and interfacial profiles shows that $CO_2$ hydration is equally as feasible under these two solvation environments and that acidification rates are likely enhanced by this additional surface contribution.},

keywords = {Chemical Reactivity, Machine Learning Potentials, Water at Interfaces},

pubstate = {published},

tppubtype = {article}

}

Yongkang Wang, Haojian Luo, Xavier R. Advincula, Zhengpu Zhao, Ali Esfandiar, Da Wu, Kara D. Fong, Lei Gao, Arsh S. Hazrah, Takashi Taniguchi, Christoph Schran, Yuki Nagata, Lydéric Bocquet, Marie-Laure Bocquet, Ying Jiang, Angelos Michaelides, Mischa Bonn
Spontaneous surface charging and Janus nature of the hexagonal boron nitride–water interface Journal Article
In: J. Am. Chem. Soc., vol. 147, iss. 33, pp. 30107–30116, 2025.
Abstract | Links | BibTeX | Tags: Charge Transfer, Machine Learning Potentials, Water at Interfaces
@article{Wang2025/10.1021/jacs.5c07827,

title = {Spontaneous surface charging and Janus nature of the hexagonal boron nitride–water interface},

author = {Yongkang Wang and Haojian Luo and Xavier R. Advincula and Zhengpu Zhao and Ali Esfandiar and Da Wu and Kara D. Fong and Lei Gao and Arsh S. Hazrah and Takashi Taniguchi and Christoph Schran and Yuki Nagata and Lydéric Bocquet and Marie-Laure Bocquet and Ying Jiang and Angelos Michaelides and Mischa Bonn},

url = {https://pubs.acs.org/doi/full/10.1021/jacs.5c07827},

doi = {10.1021/jacs.5c07827},

year  = {2025},

date = {2025-08-06},

urldate = {2025-08-06},

journal = {J. Am. Chem. Soc.},

volume = {147},

issue = {33},

pages = {30107–30116},

abstract = {Boron, nitrogen, and carbon are neighbors in the periodic table and can form strikingly similar twin structures─hexagonal boron nitride (hBN) and graphene─yet nanofluidic experiments demonstrate drastically different water friction on them. We investigate this discrepancy by probing the interfacial water and atomic-scale properties of hBN using surface-specific vibrational spectroscopy, atomic-resolution atomic force microscopy (AFM), and machine learning-based molecular dynamics. Spectroscopy reveals that pristine hBN acquires significant negative charges upon contacting water at neutral pH, unlike hydrophobic graphene, leading to interfacial water alignment and stronger hydrogen bonding. AFM supports that this charging is not defect-induced. pH-dependent measurements suggest $OH^–$ chemisorption and physisorption, which simulations validate as two nearly equally stable states undergoing dynamic exchange. These findings challenge the notion of hBN as chemically inert and hydrophobic, revealing its spontaneous surface charging and Janus nature, and providing molecular insights into its higher water friction compared to carbon surfaces.},

keywords = {Charge Transfer, Machine Learning Potentials, Water at Interfaces},

pubstate = {published},

tppubtype = {article}

}
Xavier R. Advincula, Kara D. Fong, Angelos Michaelides, Christoph Schran
Protons accumulate at the graphene–water interface Journal Article
In: ACS Nano, vol. 19, iss. 18, pp. 17728–17737, 2025.
Abstract | Links | BibTeX | Tags: Hydrogen Bonding, Machine Learning Potentials, Water, Water at Interfaces
@article{Advincula2025/10.1021/acsnano.5c02053,

title = {Protons accumulate at the graphene–water interface},

author = {Xavier R. Advincula and Kara D. Fong and Angelos Michaelides and Christoph Schran},

url = {https://pubs.acs.org/doi/full/10.1021/acsnano.5c02053},

doi = {10.1021/acsnano.5c02053},

year  = {2025},

date = {2025-04-28},

urldate = {2025-04-28},

journal = {ACS Nano},

volume = {19},

issue = {18},

pages = {17728–17737},

abstract = {Water’s ability to autoionize into hydroxide and hydronium ions profoundly influences surface properties, rendering interfaces either basic or acidic. While it is well-established that protons show an affinity to the air–water interface, a critical knowledge gap exists in technologically relevant surfaces like the graphene–water interface. Here we use machine learning-based simulations with first-principles accuracy to unravel the behavior of hydroxide and hydronium ions at the graphene–water interface. Our findings reveal that protons accumulate at the graphene–water interface, with the hydronium ion predominantly residing in the first contact layer of water. In contrast, the hydroxide ion exhibits a bimodal distribution, found both near the surface and further away from it. Analysis of the underlying electronic structure reveals local polarization effects, resulting in counterintuitive charge rearrangement. Proton propensity to the graphene–water interface challenges the interpretation of surface experiments and is expected to have far-reaching consequences for ion conductivity, interfacial reactivity, and proton-mediated processes.},

keywords = {Hydrogen Bonding, Machine Learning Potentials, Water, Water at Interfaces},

pubstate = {published},

tppubtype = {article}

}
2024
Samuel G.H. Brookes, Venkat Kapil, Christoph Schran, Angelos Michaelides
The wetting of H₂O by CO₂ Journal Article
In: J. Chem. Phys., vol. 161, no. 8, pp. 084711, 2024, ISSN: 10897690.
Abstract | Links | BibTeX | Tags: Machine Learning Potentials, Water, Water at Interfaces
@article{Brookes2024/10.1063/5.0224230,

title = {The wetting of H₂O by CO₂},

author = {Samuel G.H. Brookes and Venkat Kapil and Christoph Schran and Angelos Michaelides},

url = {/aip/jcp/article/161/8/084711/3309975/The-wetting-of-H2O-by-CO2},

doi = {10.1063/5.0224230},

issn = {10897690},

year  = {2024},

date = {2024-08-28},

urldate = {2024-08-28},

journal = {J. Chem. Phys.},

volume = {161},

number = {8},

pages = {084711},

publisher = {American Institute of Physics},

abstract = {Biphasic interfaces are complex but fascinating regimes that display a number of properties distinct from those of the bulk. The CO2-H2O interface, in particular, has been the subject of a number of studies on account of its importance for the carbon life cycle as well as carbon capture and sequestration schemes. Despite this attention, there remain a number of open questions on the nature of the CO2-H2O interface, particularly concerning the interfacial tension and phase behavior of CO2 at the interface. In this paper, we seek to address these ambiguities using ab initio-quality simulations. Harnessing the benefits of machine-learned potentials and enhanced statistical sampling methods, we present an ab initio-level description of the CO2-H2O interface. Interfacial tensions are predicted from 1 to 500 bars and found to be in close agreement with experiment at pressures for which experimental data are available. Structural analyses indicate the buildup of an adsorbed, saturated CO2 film forming at a low pressure (20 bars) with properties similar to those of the bulk liquid, but preferential perpendicular alignment with respect to the interface. The CO2 monolayer buildup coincides with a reduced structuring of water molecules close to the interface. This study highlights the predictive nature of machine-learned potentials for complex macroscopic properties of biphasic interfaces, and the mechanistic insight obtained into carbon dioxide aggregation at the water interface is of high relevance for geoscience, climate research, and materials science.},

keywords = {Machine Learning Potentials, Water, Water at Interfaces},

pubstate = {published},

tppubtype = {article}

}
2022
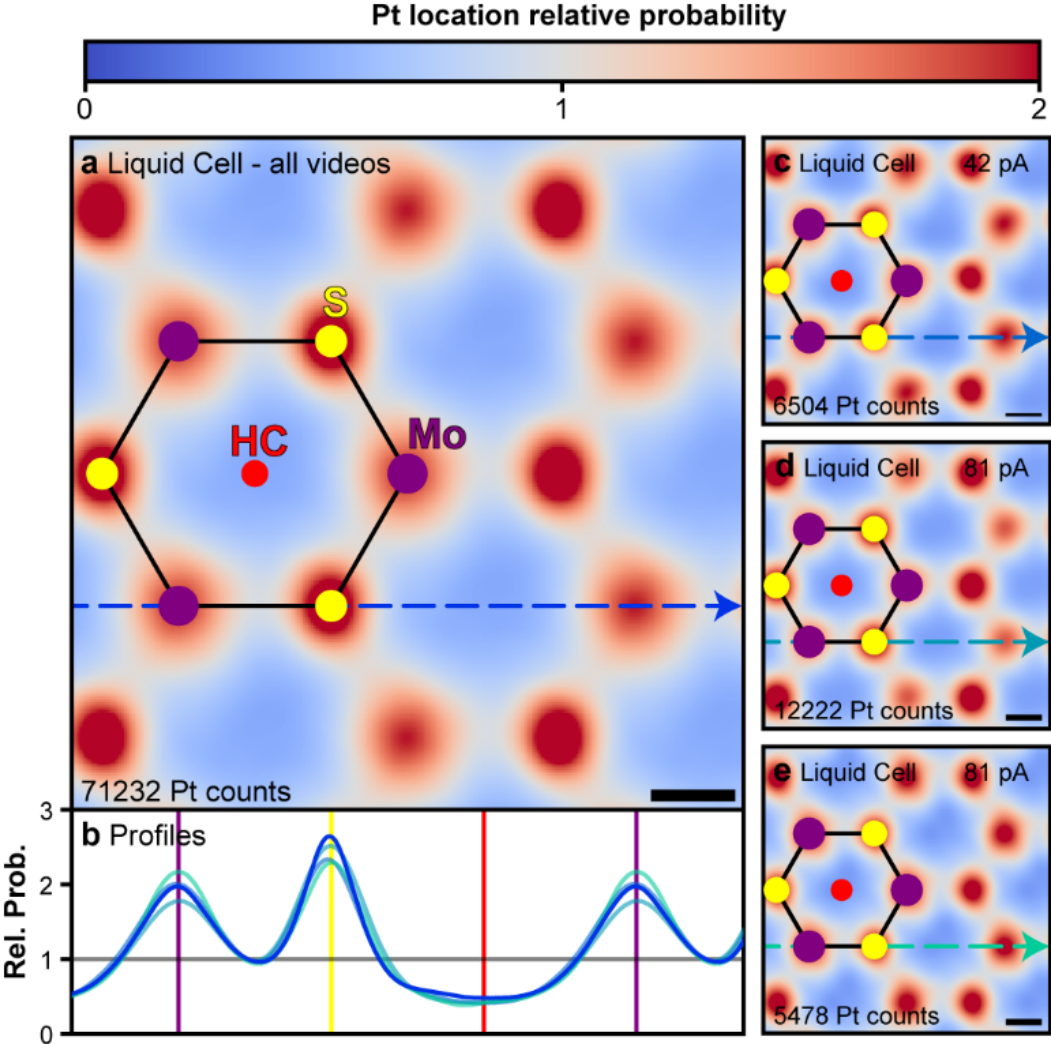
Nick Clark, Daniel J. Kelly, Mingwei Zhou, Yi-Chao Zou, Chang Woo Myung, David G. Hopkinson, Christoph Schran, Angelos Michaelides, Roman Gorbachev, Sarah J. Haigh
Tracking single adatoms in liquid in a Transmission Electron Microscope Journal Article
In: Nature, pp. 1–3, 2022, ISSN: 1476-4687.
Abstract | Links | BibTeX | Tags: Two-dimensional Materials, Water at Interfaces
@article{Clark2022/10.1038/s41586-022-05130-0,

title = {Tracking single adatoms in liquid in a Transmission Electron Microscope},

author = {Nick Clark and Daniel J. Kelly and Mingwei Zhou and Yi-Chao Zou and Chang Woo Myung and David G. Hopkinson and Christoph Schran and Angelos Michaelides and Roman Gorbachev and Sarah J. Haigh},

doi = {10.1038/s41586-022-05130-0},

issn = {1476-4687},

year  = {2022},

date = {2022-07-01},

urldate = {2022-07-01},

journal = {Nature},

pages = {1–3},

publisher = {Nature Publishing Group},

abstract = {Single atoms or ions on surfaces affect processes from nucleation1 to electrochemical reactions2 and heterogeneous catalysis3. Transmission electron microscopy (TEM) is a leading approach for visualizing single atoms on a variety of substrates4,5. It conventionally requires high vacuum conditions, but has been developed for in situ imaging in liquid and gaseous environments6,7 with a combined spatial and temporal resolution that is unmatched by any other method —notwithstanding concerns about electron beam effects on samples. When imaging in liquid using commercial technologies, electron scattering in the windows enclosing the sample and in the liquid generally limits the achievable resolution to a few nanometres6,8,9. Graphene liquid cells, on the other hand, have enabled atomic resolution imaging of metal nanoparticles in liquids10. Here we show that a double graphene liquid cell, comprised of a central molybdenum disulphide monolayer separated by hexagonal boron nitride spacers from the two enclosing graphene windows, makes it possible to monitor with atomic resolution the dynamics of platinum adatoms on the monolayer in an aqueous salt solution. By imaging over 70,000 single adatom adsorption sites, we compare the site preference and dynamic motion of the adatoms in both a fully hydrated and vacuum state. We find a modified adsorption site distribution and higher diffusivities for the adatoms in liquid phase compared to those in vacuum. This approach paves the way for in situ liquid phase imaging of chemical processes with single atom precision.},

keywords = {Two-dimensional Materials, Water at Interfaces},

pubstate = {published},

tppubtype = {article}

}
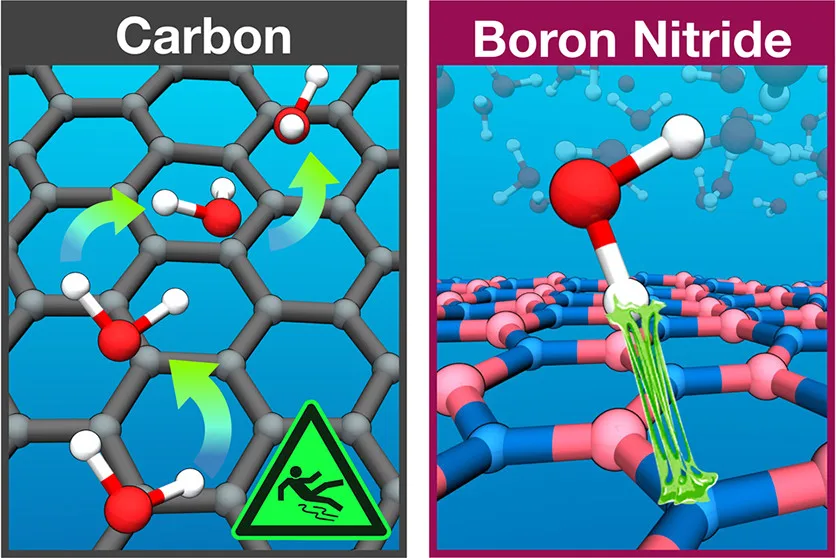
Fabian L. Thiemann, Christoph Schran, Patrick Rowe, Erich A. Müller, Angelos Michaelides
Water flow in single-wall nanotubes: Oxygen makes it slip, hydrogen makes it stick Journal Article
In: ACS Nano, vol. 16, no. 7, pp. 10775–10782, 2022.
Abstract | Links | BibTeX | Tags: Confinement, Hydrogen Bonding, Machine Learning Potentials, Water at Interfaces, Water Flow
@article{Thiemann2022/10.1021/acsnano.2c02784,

title = {Water flow in single-wall nanotubes: Oxygen makes it slip, hydrogen makes it stick},

author = {Fabian L. Thiemann and Christoph Schran and Patrick Rowe and Erich A. Müller and Angelos Michaelides},

doi = {10.1021/acsnano.2c02784},

year  = {2022},

date = {2022-06-01},

urldate = {2022-06-01},

journal = {ACS Nano},

volume = {16},

number = {7},

pages = {10775–10782},

abstract = {Experimental measurements have reported ultra-fast and radius-dependent water transport in carbon nanotubes which are absent in boron nitride nanotubes. Despite considerable effort, the origin of this contrasting (and fascinating) behaviour is not understood. Here, with the aid of machine learning-based molecular dynamics simulations that deliver first-principles accuracy, we investigate water transport in single-wall carbon and boron nitride nanotubes. Our simulations reveal a large, radius-dependent hydrodynamic slippage on both materials with water experiencing indeed a $backslashapprox 5$ times lower friction on carbon surfaces compared to boron nitride. Analysis of the diffusion mechanisms across the two materials reveals that the fast water transport on carbon is governed by facile oxygen motion, whereas the higher friction on boron nitride arises from specific hydrogen-nitrogen interactions. This work not only delivers a clear reference of unprecedented accuracy for water flow in single-wall nanotubes, but also provides detailed mechanistic insight into its radius and material dependence for future technological application.},

keywords = {Confinement, Hydrogen Bonding, Machine Learning Potentials, Water at Interfaces, Water Flow},

pubstate = {published},

tppubtype = {article}

}
2021
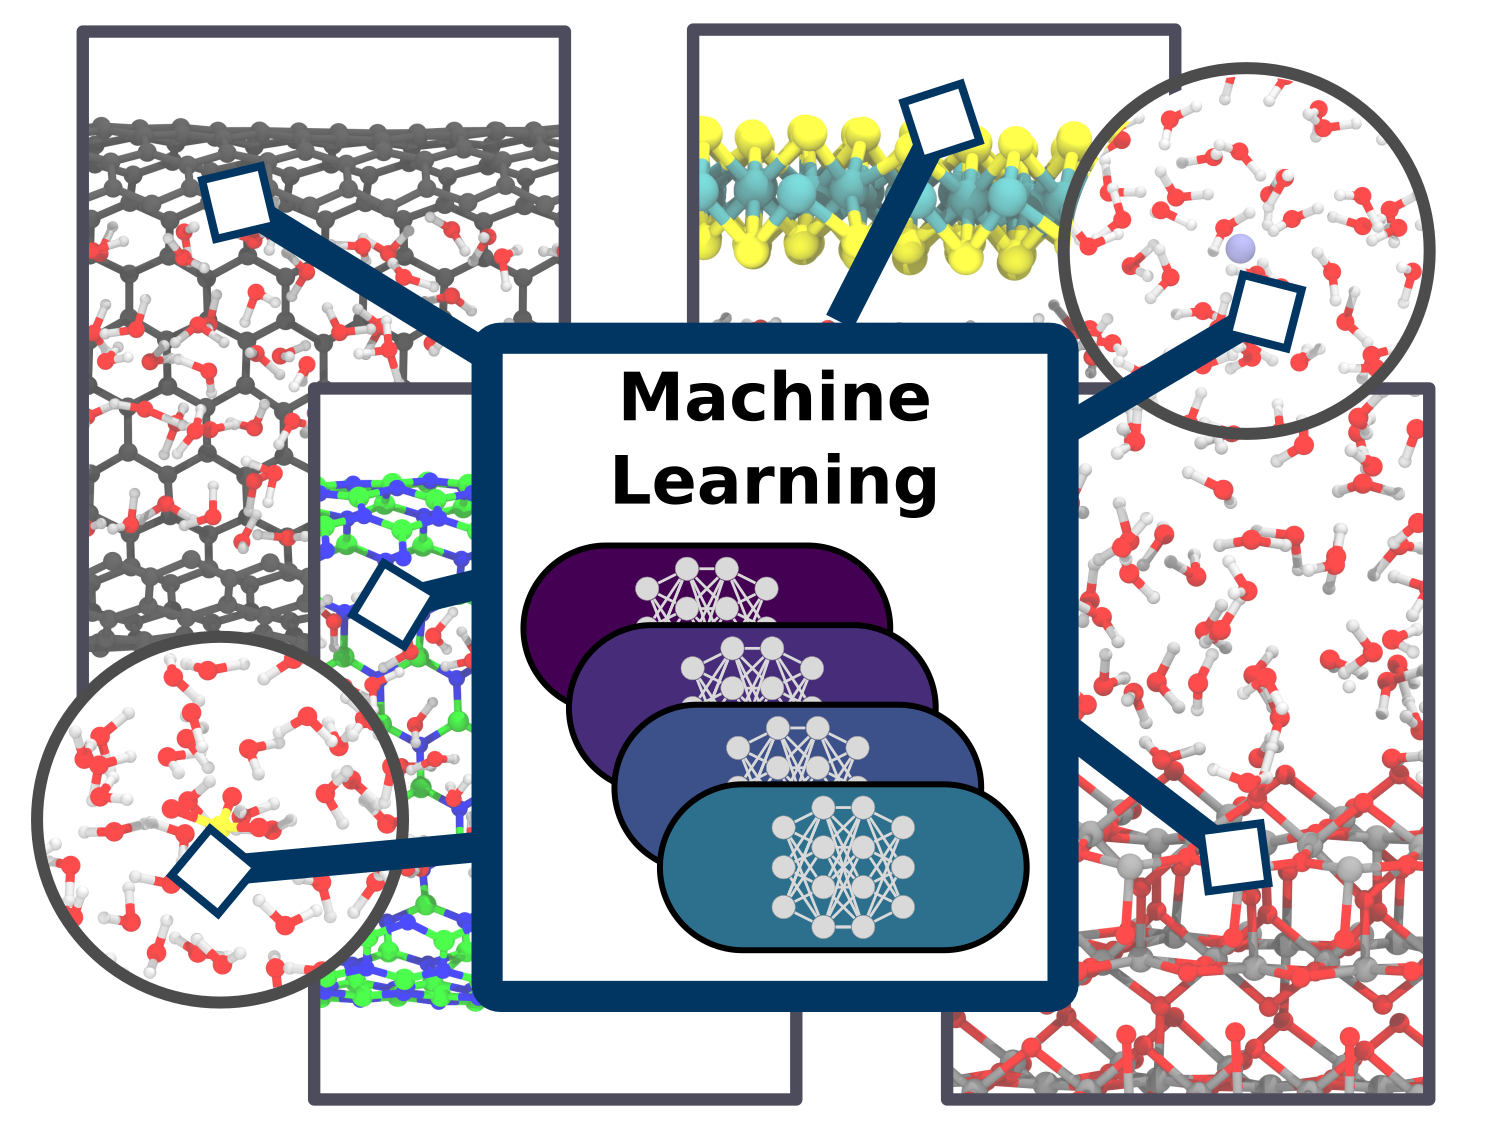
Christoph Schran, Fabian L. Thiemann, Patrick Rowe, Erich A. Müller, Ondrej Marsalek, Angelos Michaelides
Machine learning potentials for complex aqueous systems made simple Journal Article
In: Proc. Natl. Acad. Sci., vol. 118, no. 38, pp. e2110077118, 2021, ISSN: 0027-8424.
Abstract | Links | BibTeX | Tags: Confinement, Ions in Water, Machine Learning Potentials, Water, Water at Interfaces
@article{Schran2021/10.1073/PNAS.2110077118,

title = {Machine learning potentials for complex aqueous systems made simple},

author = {Christoph Schran and Fabian L. Thiemann and Patrick Rowe and Erich A. Müller and Ondrej Marsalek and Angelos Michaelides},

doi = {10.1073/PNAS.2110077118},

issn = {0027-8424},

year  = {2021},

date = {2021-09-01},

urldate = {2021-09-01},

journal = {Proc. Natl. Acad. Sci.},

volume = {118},

number = {38},

pages = {e2110077118},

publisher = {National Academy of Sciences},

abstract = {Simulation techniques based on accurate and efficient representations of potential energy surfaces are urgently needed for the understanding of complex aqueous systems such as solid-liquid interfaces. Here, we present a machine learning framework that enables the efficient development and validation of models for complex aqueous systems. Instead of trying to deliver a globally-optimal machine learning potential, we propose to develop models applicable to specific thermodynamic state points in a simple and user-friendly process. After an initial ab initio simulation, a machine learning potential is constructed with minimum human effort through a data-driven active learning protocol. Such models can afterwards be applied in exhaustive simulations to provide reliable answers for the scientific question at hand. We showcase this methodology on a diverse set of aqueous systems with increasing degrees of complexity. The systems chosen here comprise water with different ions in solution, water on a titanium dioxide surface, as well as water confined in nanotubes and between molybdenum disulfide sheets. Highlighting the accuracy of our approach compared to the ab initio reference, the resulting models are evaluated in detail with an automated validation protocol that includes structural and dynamical properties and the precision of the force prediction of the models. Finally, we demonstrate the capabilities of our approach for the description of water on the rutile titanium dioxide (110) surface to analyze structure and mobility of water on this surface. Such machine learning models provide a straightforward and uncomplicated but accurate extension of simulation time and length scales for complex systems.},

keywords = {Confinement, Ions in Water, Machine Learning Potentials, Water, Water at Interfaces},

pubstate = {published},

tppubtype = {article}

}